X THE PROGRESS OF CHEMISTRY DURING THE PAST ONE HUNDRED YEARS
Reading edition
By HORACE L. WELLS and HARRY W. FOOTE
Introduction.
As we look back to the time of the founding of the Journal in 1818, we see that the science of chemistry had recently made and was then making great advances. That the scientific men of those days were much impressed with what was being accomplished is well shown by the following statement made in an early number of the Journal (3, 330, 1821) by its founder in reviewing Gorham’s Elements of Chemical Science. He says: “The present period is distinguished by wonderful mental activity; it might indeed be denominated as the intellectual age of the world. At no former period has the mind of man been directed at one time to so many and so useful researches.”
A very remarkable revolution in chemical ideas had recently taken place. Soon after the discovery of oxygen by Priestley in 1774, and the subsequent discovery by Cavendish that water was formed by the combustion of hydrogen and oxygen, Lavoisier had explained combustion in general as oxidation, thus overthrowing the curious old phlogiston theory which had prevailed as the basis of chemical philosophy for nearly a century.
The era of modern chemistry had thus begun, and the additional views that matter was indestructible and that chemical compounds were of constant composition had been generally accepted at the beginning of the nineteenth century.
Dalton had announced his atomic theory in 1802, having based it largely upon the law of multiple proportions 289which he had previously discovered, and he had begun to express the formulas for compounds in terms of atomic symbols.
In 1808 Gay-Lussac had discovered his law of gas combination in simple proportions,[153] a law of supreme importance in connection with the atomic theory, but neither he nor Dalton had seen this theoretical connection. Avogadro had understood it, however, and in 1811 had reached the momentous conclusion that all gases and vapors have equal numbers of molecules in equal volumes at the same temperature and pressure.
Davy in 1807 had isolated the alkali-metals, sodium and potassium, by means of electrolysis, thus practically dispelling the view that certain earthy substances might be elementary; and about four years later he had demonstrated that chlorine was an element, not an oxide as had been supposed previously, thus overthrowing Lavoisier’s view that oxygen was the characteristic constituent of all acids.
At the time that our period of history begins, the atomic theory had been accepted generally, but in a somewhat indefinite form, since little attention had been paid to Avogadro’s principle, and since Dalton had used only the principle of greatest simplicity in writing the formulas of compounds, considering water as HO and ammonia NH, for example. At this time, however, Berzelius for ten or fifteen years had been devoting tremendous energy to the task of determining the atomic weights of nearly all of the elements then known by analyzing their compounds. He had confirmed the law of multiple proportions, accepted the atomic theory, and utilized Avogadro’s principle, and it is an interesting coincidence that his first table of atomic weights was published in the year 1818.
An interesting account of the views on chemistry held at about that time was published in the Journal by Denison Olmsted (11, 349, 1826; 12, 1, 1827), who had recently become professor of natural philosophy in Yale College.
The most illustrious European chemists of that time were Berzelius of Sweden, Davy of England, and Gay-Lussac of France, and the curious circumstance may be 290mentioned that all three of them and also Benjamin Silliman, the founder of the Journal, were born within a period of eight months in 1778–1779.
In this country Robert Hare of Philadelphia and Benjamin Silliman were undoubtedly the most prominent chemists of those days. Hare is best known for his invention of the compound blowpipe, but his contributions to the Journal were very numerous, beginning almost with the first volume and continuing for over thirty years. Among the first of these contributions was a most vigorous but well-merited attack upon a Doctor Clark of Cambridge, England, who had copied his invention without giving him proper credit. He begins (2, 281, 1820) by saying: “Dr. Clark has published a book on the gas blowpipe in which he professes a sincere desire to render everyone his due. That it would be difficult for the conduct of any author to be more discordant with these professions, I pledge myself to prove in the following pages.”
Hare also invented a galvanic battery which he called a “deflagrator,” consisting of a large number of single cells in series. With this, using carbon electrodes, he was able to obtain a higher temperature than with his oxy-hydrogen blowpipe. He was the first to apply galvanic ignition to blasting (21, 139, 1832), and he first carried out electrolyses with the use of mercury as the cathode (37, 267, 1839). In this way he prepared metallic calcium and other metals from solutions of their chlorides, while the principle employed by him has in recent times been used as the basis of a very important process for manufacturing caustic potash and soda.
Silliman, who had become an intimate friend of Hare during two periods of chemical study under Woodhouse in Philadelphia in 1802–1804, and who soon afterwards spent fourteen months as a student abroad, chiefly in England and Scotland, took a broad interest in science and gave much attention to geology as well as to chemistry. In spite of this divided interest and his work as a teacher, popular scientific lecturer, and editor, he found time for a surprising amount of original chemical work. For instance, using Hare’s deflagrator, he showed that carbon was volatilized in the electric arc (5, 108, 1822); 291he was the first in this country to prepare hydrofluoric acid (6, 354, 1823), and he first detected bromine in one of our natural brines (18, 142, 1830).
Atomic Weights.
As soon as the atomic theory was accepted, the relative weights of the atoms became a matter of vital importance in connection with formulas and chemical calculations. In advancing his theory, Dalton had made some very rough atomic weight determinations, and it has been mentioned already that Berzelius, at the time that our historical period begins, was engaged in the prodigious task of accurately determining these constants for nearly all the known elements. It is recorded that he analyzed quantitatively no less than two thousand compounds in connection with this work during his career. His table of 1818 has proved to be remarkably accurate for that pioneer period, and it indicates his remarkable skill as an analyst.
It is to be observed that Berzelius in this early table made use of Avogadro’s principle in connection with elements forming gaseous compounds, and thus obtained correct formulas and atomic weights in such cases, but that in many instances his atomic weights and those now accepted bear the relation of simple multiples to one another, because he had then no means of deciding upon the formulas of many compounds except the rule of assumed simplicity. For example, the two oxides of iron now considered to be FeO and Fe2O3 he regarded as FeO2 and FeO3, knowing as he did that the ratio of oxygen in them was 2 to 3, and believing that a single atom of iron in each was the simplest view of the case, so that as the consequence of these formulas the atomic weight of iron was then considered to be practically twice as great in its relation to oxygen as at present.
These old atomic weights of Berzelius, used with the corresponding formulas, were just as serviceable for calculating compositions and analytical factors as though the correct multiples had been selected. As time went on, the true multiples were gradually found from considerations of atomic heats, isomorphism, vapor densities, 292the periodic law, and so on, and suitable changes were made in the chemical formulas.
Berzelius used 100 parts of oxygen as the basis of his atomic weights, a practice which was generally followed for several decades. Dalton, however, had originally used hydrogen as unity as the basis, and this plan finally came into use everywhere, as it seemed to be more logical and convenient, because hydrogen has the smallest atomic weight, and also because the atomic weights of a number of common elements appeared to be exact multiples of that of hydrogen, thus giving simpler numbers for use in calculations.
Within a few years a slight change has been made by the adoption of oxygen as exactly 16 as the basis, which gives hydrogen the value of 1·008.
As early as 1815, Prout, an English physician, had advanced the view that hydrogen is the primordial substance of all the elements, and consequently that the atomic weights are all exact multiples of that of hydrogen. This hypothesis has been one of the incentives to investigations upon atomic weights, for it has been found that these constants in the cases of a considerable number of the elements are very close to whole numbers when based upon hydrogen as unity, or even still closer when based upon oxygen as 16.
With our present knowledge Prout’s hypothesis may be regarded as disproved for nearly all the elements whose atomic weights have been accurately determined, but the close or even exact agreement with it in a few cases is still worthy of consideration. There is an interesting letter from Berzelius to B. Silliman, Jr., in the Journal (48, 369, 1845) in which Berzelius considers the theory entirely disproved.
For a long time entire reliance was placed upon the atomic weights obtained by Berzelius, but it came to be observed that the calculation of carbon from carbon dioxide appeared to give high results in certain cases, so that doubt arose as to the accuracy of Berzelius’s work. Consequently in 1840 Dumas, assisted by his pupil Stas, made a new determination of the atomic weight of carbon, and found that the number obtained by Berzelius, 12·12, was slightly too large. Subsequently Dumas determined 293more than twenty other atomic weights, but this great amount of work did not bring about any considerable improvement, for it appears that Dumas did not greatly excel Berzelius in accuracy, and that the latter had made one of his most noticeable errors in connection with carbon.
Soon after assisting Dumas in the work upon carbon, Stas began his very extensive and accurate, independent determinations, leading to the publication of a book in 1867 describing his work. Stas made many improvements in methods by the use of great care in purifying the substances employed, and especially by using large quantities of material in his determinations, thus diminishing the proportional errors in weighing. His results, which dealt with most of the common elements, were accepted with much confidence by chemists everywhere.
Stas reached the conclusion that there could be no real foundation for Prout’s hypothesis, since so many of his atomic weights varied from whole numbers, and this opinion has been generally accepted.
The first accurate atomic weight determination published in the Journal was that by Mallett on lithium (22, 349, 1856; 28, 349, 1859), showing a result almost identical with that accepted at the present time. Johnson and Allen’s determination (35, 94, 1863) on the rare element cæsium was carried out with extraordinary accuracy. Lee, working with Wolcott Gibbs, made good determinations on nickel and cobalt (2, 44, 1871). The work of Cooke on antimony (15, 41, 107, 1878) was excellent.
Concerning the more recent work published elsewhere than in the Journal, attention should be called particularly to the investigations that have been carried on for the past twenty-five years by Richards and his associates at Harvard University. Richards has shown masterly ability in the selection of methods and in avoiding errors. His results have displayed such marvelous agreements among repeated determinations by the same and by different processes as to inspire the greatest confidence. His work has been very extensive, and it is a great credit to our country that this atomic weight work, so superior to all that has been previously done, is being carried out here.
294It may be mentioned that for a number of years the decision in regard to the atomic weights to be accepted has been in the hands of an International Committee of which our fellow countryman F. W. Clarke has been chairman. In connection with this position and previously, Clarke has done valuable service in re-calculating and summarizing atomic weight determinations.
Analytical Chemistry.
Analysis is of such fundamental importance in nearly every other branch of chemical investigation that its development has been of the utmost importance in connection with the advancement of the science. It attained, therefore, a comparatively early development, and one hundred years ago it was in a flourishing condition, particularly as far as inorganic qualitative and gravimetric analysis were concerned. There is no doubt that Berzelius, whose atomic weight determinations have already been mentioned, surpassed all other analysts of that time in the amount, variety, and accuracy of his gravimetric work. He lived through three decades of our period, until 1848.
During the past century there has been constant progress in inorganic analysis, due to improved methods, better apparatus and accumulated experience. An excellent work on this subject was published by H. Rose, a pupil of Berzelius, and the methods of the latter, with many improvements and additions by the author and others, were thus made accessible. Fresenius, who was born in 1818, did much service in establishing a laboratory in which the teaching of analytical chemistry was made a specialty, in writing text-books on the subject and in establishing in 1862 the “Zeitschrift für analytische Chemie,” which has continued up to the present time.
Besides Berzelius, who was the first to show that minerals were definite chemical compounds, there have been many prominent mineral analysts in Europe, among whom Rammelsberg and Bunsen may be mentioned, but there came a time towards the end of the nineteenth century when the attention of chemists, particularly in Germany, was so much absorbed by organic chemistry that 295mineral analysis came near becoming a lost art there. It was during that period that an English mineralogist, visiting New Haven and praising the mineral analyses that were being carried out at Yale, expressed regret that there appeared to be no one in England, or in Germany either, who could analyze minerals.
The best analytical work done in this country in the early part of our period was chiefly in connection with mineral analysis, and a large share of it was published in the Journal. Henry Seybert, of Philadelphia, in particular, showed remarkable skill in this direction, and published numerous analyses of silicates and other minerals, beginning in 1822. It was he who first detected boric acid in tourmaline (6, 155, 1822), and beryllium in chrysoberyl (8, 105, 1824). His methods for silicate analyses were very similar to those used at the present time.
J. Lawrence Smith in 1853 described his method for determining alkalies in minerals (16, 53), a method which in its final form (1, 269, 1871) is the best ever devised for the purpose. He also described (15, 94, 1853) a very useful method, still largely used in analytical work, for destroying ammonium salts by means of aqua regia. Carey Lea (42, 109, 1866) described the well-known test for iodides by means of potassium dichromate. F. W. Clarke (49, 48, 1870) showed that antimony and arsenic could be quantitatively separated from tin by the precipitation of the sulphides in the presence of oxalic acid. In 1864 Wolcott Gibbs (37, 346) began an important series of analytical notes from the Lawrence Scientific School, and he worked out later many difficult analytical problems, particularly in connection with his extensive researches upon the complex inorganic acids.
From 1850 on, Brush and his students made many important investigations upon minerals, and from 1877 Penfield (13, 425), beginning with an analysis of a new mineral from Branchville, Connecticut, described by Brush and E. S. Dana, displayed remarkable skill and industry in this kind of work. Both of the writers of this article were fortunate in being associated with Penfield in some of his researches upon minerals and one of us began as he did with the Branchville work. It is 296probably fair to say that Penfield did the most accurate work in mineral analysis that has ever been accomplished, and that he was similarly successful in crystallography and other physical branches of mineralogy.
The American analytical investigations that have been mentioned were all published in the Journal, with the exception of a part of Gibbs’s work. Many other American workers at mineral analysis might be alluded to here, but only the excellent work of a number of chemists in the United States Geological Survey will be mentioned. Among these Hillebrand deserves particular praise for the extent of his investigations and for his careful researches in improving the methods of rock analysis.
To our own Professor Gooch especial praise must be accorded for the very large number of analytical methods that have been devised, or critically studied, by him and his students, and for the excellent quality of this work. The publications in the Journal from his laboratory began in 1890 (39, 188), and the extraordinary extent of this work is shown by the fact that the three hundredth paper from the Kent Laboratory appeared in May, 1918. These very numerous and important investigations have been of great scientific and practical value, and they have formed a striking feature of the Journal for nearly 30 years. In 1912 Gooch published his “Methods in Chemical Analysis,” a book of over 500 pages, in which the work in the Kent Chemical Laboratory up to that time was concisely presented. Among the many workers who have assisted in these investigations, P. E. Browning, W. A. Drushel, F. S. Havens, D. A. Kreider, C. A. Peters, I. K. Phelps and R. G. Van Name are particularly prominent. Besides many other useful pieces of apparatus, the perforated filtering crucible was devised by Gooch, and this has brought his name into everyday use in all chemical laboratories.
Volumetric analysis was originated by Gay-Lussac, who described a method for chlorimetry in 1824, for alkalimetry in 1828, and for the determination of silver and chlorides in 1832. Margueritte devised titrations with potassium permanganate in 1846, while Bunsen, not far from the same time, introduced the use of iodine and sulphur dioxide solutions for the purpose of determining 297many oxidations and reductions. We owe to Mohr some improvements in apparatus and a German text-book on the subject, while Sutton wrote an excellent English work on volumetric analysis, of which many editions have appeared.
While volumetric analysis began to be used less than one hundred years ago, its applications have been gradually extended to a very great degree, and it is not only exceedingly important in investigations in pure chemistry, but its use is especially extensive in technical laboratories where large numbers of rapid analyses are required.
Not a few volumetric methods have been devised or improved in the United States, but mention will be made here only of Cooke’s important method for the determination of ferrous iron in insoluble silicates, published in the Journal (44, 347, 1867); to Penfield’s method for the determination of fluorine in 1878; and to the more recent general method of titration with an iodate in strong hydrochloric acid solutions, due to L. W. Andrews, a number of applications of which have been worked out in the Sheffield Laboratory.
A considerable amount of work with gases had been done by Priestley, Scheele, Cavendish, Lavoisier, Dalton, Gay-Lussac, and others before our hundred-year period began. Cavendish, about 1780, had analyzed atmospheric air with remarkable accuracy, and had even separated the argon from it and wondered what it was, and later Gay-Lussac had shown great skill in the study of gas reactions. During our period gas analysis has been further developed by many chemists. Bunsen, in particular, brought the art to a high degree of perfection in the course of a long period beginning about 1838, the last edition of his “Methods of Gas Analysis” having been published in 1877.
Important devices for the simplification of gas analysis in order that it might be used more conveniently for technical purposes have been introduced by Orsat in France and by Winkler, Hempel and Bunte in Germany.
It appears that our countryman Morley has surpassed all others in accurate work with gases in connection with his determinations of the combining weights and 298volumes of hydrogen and oxygen about the year 1891. Some of his publications have appeared in the Journal (30, 140, 1885; 41, 220, 1891; and others).
Electrolytic analysis, involving the deposition of metals, or sometimes of oxides, usually upon a platinum electrode, was brought into use in 1865 by Wolcott Gibbs through an article published in the Journal (39, 58, 1865). He there described the electrolytic precipitation of copper and of nickel by the methods still in use. The application of the process has been extended to a number of other metals, and it has been largely employed, particularly in technical analyses. Important investigations and excellent books on this subject have been the contributions of Edgar F. Smith of the University of Pennsylvania, and the useful improvement, the rotating cathode, was devised by Gooch and described in the Journal (15, 320, 1903).
General Inorganic Chemistry.
The Chemical Symbols.—It is to Berzelius that we owe our symbols for the atoms, derived usually from their Latin names, such as C for carbon, Na for sodium, Cl for chlorine, Fe for iron, Ag for silver, and Au for gold. We owe to him also the use of small figures to show the number of atoms in a formula, as in N2O5. This was a marked improvement over the hieroglyphic symbols proposed by Dalton, which were set down as many times as the atoms were supposed to occur in formulas, forming groups of curious appearance, but in some respects not unlike some of our modern developed formulas. The advantages of Berzelius’s symbols were their simplicity, legibility, and the fact that they could be printed without the need of special type. It is true that at a later period Berzelius used certain symbols with horizontal lines crossing them to represent double atoms, and that these made some difficulty in printing. It should be mentioned also that Berzelius at one time made an effort to simplify formulas by placing dots over other symbols to represent oxygen, and commas to represent sulphur atoms. Examples of these are:
ĊaS⃛, calcium sulphate; F̋e, iron disulphide
299This form of notation was quite extensively employed for a time, especially by mineralogists, but it was entirely abandoned later.
It is interesting to notice that Dalton, who lived until 1844, to reach the age of 78, differed from other chemists in refusing to accept the letter-symbols of Berzelius. In a letter written to Graham in 1837 he said: “Berzelius’s symbols are horrifying. A young student in chemistry might as soon learn Hebrew as to make himself acquainted with them. They appear like a chaos of atoms ... and to equally perplex the adepts of science, to discourage the learner, as well as to cloud the beauty and simplicity of the atomic theory.”
This forcibly expressed opinion was apparently tinged with self-esteem, but there is no doubt that Dalton was sincere in believing that the atoms were best represented by his circular symbols, because, as is well known, he thought that all the atoms were spherical in form, and it is evident that circles give the proper picture of spherical objects. At the present time some insight as to the structure of atoms is being gained, and it appears possible that the time may come when pictures of their external appearance that are not wholly imaginary may be made.
Changes in Formulas.—Even before the year 1826, Berzelius displayed great skill in arriving at many formulas that agree with our present ones, for example, H2O for water, ZnCl2 for zinc chloride, N2O5 for nitric acid (anhydride), CaO for calcium oxide, CO and CO2 for the oxides of carbon, and many others. But at the same period other authorities, especially Gay-Lussac in France and Gmelin in Germany, on account of a lack of appreciation for Avogadro’s principle and for other reasons, such as the use of symbols to represent combining weights rather than atoms, were using different formulas for some of these compounds, such as HO, ZnCl and NO5, so that their formulas for many of the compounds of hydrogen, chlorine, nitrogen and several other elements differed from those of Berzelius. The employment of different formulas involved the use of different atomic or combining weights. For example, with the formula H2O for water the composition by weight requires the 300ratio 1 to 16 for the weights of the hydrogen and oxygen atoms, while with HO the ratio is 1 to 8.
Berzelius attempted to bring about greater uniformity in formulas and atomic weights by making changes in his table of atomic weights published in 1826. He practically doubled the relative atomic weights of hydrogen, chlorine, nitrogen, and of the other elements that gave twice as many atoms in his formulas as in those of others, and at the same time he wrote the symbols of these elements with a bar across them to indicate that they represented double atoms. For example, he wrote:
H̶O ZnC̶l N̶O5,
instead of
H2O, ZnCl2 N2O5
This appears to have been an unfortunate concession to the views of others on the part of Berzelius, for the barred symbols were not generally adopted, partly on account of difficulties in printing, and the great achievement in theory made by him was lost sight of for a long period of time.
The Law of Atomic Heats.—In 1819, Dulong and Petit of France, from experiments upon the specific heats of a number of solid elementary substances, came to the conclusion that the atoms of simple substances have equal capacities for heat, or in other words, that the specific heats of elements multiplied by their atomic weights give a constant called the atomic heat. For instance, the specific heats of sulphur, iron, and gold have been given as 0·2026, 0·110, and 0·0324, while their atomic weights are about 32, 56, and 197, respectively; hence the atomic heats obtained by multiplication are 6·483, 6·116, and 6·383.
Further investigations showed that the atomic heats display a considerable variation. Those of carbon, boron, beryllium, and silicon are very low at ordinary temperatures, although they increase and approach the usual values at higher temperatures. More recent work has shown, however, that the specific heats of other elements vary greatly with the temperature, almost disappearing at the temperature of liquid hydrogen, and hence possibly disappearing entirely at the absolute zero, where 301the electrical resistance of the metals appears to vanish likewise.
It has been found that most of the solid elements near ordinary temperatures give atomic heats that are approximately 6·4. Berzelius applied the law in fixing a number of atomic weights, and its importance for this purpose is still recognized.
It may be mentioned here that two well-known Yale men, W. O. Mixter and E. S. Dana, while students in Bunsen’s laboratory at Heidelberg in 1873, made determinations of the specific heats of boron, silicon, and zirconium. This was the first determination of this constant for zirconium, and it was consequently important in establishing the atomic weight of that element.
Isomorphism and Polymorphism.—Mitscherlich observed in 1818 that certain phosphates and arsenates have the same crystalline form, and afterwards he reached the conclusion that identity in form indicates similarity in composition in connection with the number of atoms and their arrangement. This law of isomorphism was of much assistance in the establishment of correct formulas and consequently of atomic weights. For instance, since the carbonates of barium, strontium, and lead crystallize in the same form, the oxides of these metals must have analogous formulas. From such considerations Berzelius was able to make several improvements in his atomic weight table of 1826.
Mitscherlich was the first to observe two forms of sulphur crystals, and from this and other cases of dimorphism or of polymorphism it became evident that analogous compounds were not necessarily always isomorphous, a circumstance which has restricted the application of the law to some extent.
Besides its application in fixing analogous formulas, the law of isomorphism has come to be of much practical use in the understanding and simplification of the formulas for minerals, for these natural crystals very often contain several isomorphous compounds in varying proportions, and an understanding of this “isomorphous replacement,” as it is called, makes it possible to deduce simple general formulas for them.
302In some cases isomorphism takes place to a greater or less extent between substances which are not chemically similar, and this brings about a variation in composition which at times has caused confusion. For instance, the mineral pyrrhotite has a composition which usually varies between Fe7S8 and Fe11S12, and both these formulas have been assigned to it. It was recently shown by Allen, Crenshaw and Johnston in the Journal (33, 169, 1912) that this is a case where the compound FeS is capable of taking up various amounts of sulphur isomorphously.
The idea of solid solution was advanced by van’t Hoff to explain the crystallization of mixtures, including cases of evident isomorphism. This view has been widely accepted, and it has been particularly useful in cases where isomorphism is not evident. Solid solution between metals has been found to be exceedingly common, many alloys being of this character. A case of this kind was observed by Cooke and described in the Journal (20, 222, 1855). He prepared two well-crystallized compounds of zinc and antimony to which he gave the formulas Zn3Sb and Zn2Sb, but he observed that excellent crystals of each could be obtained which varied largely in composition from these formulas. As the two compounds were dissimilar in their formulas and crystalline forms, Cooke assumed that isomorphism was impossible and concluded “that it is due to an actual perturbation of the law of definite proportions, produced by the influence of mass.” We should now regard this as a case of solid solution.
A Lack of Confidence in Avogadro’s Principle.—One reason why chemists were so slow in arriving at the correct atomic weights and formulas was a partial loss of confidence in Avogadro’s principle. About 1826 the young French chemist Dumas devised an excellent method for the determination of vapor densities at high temperatures, and his results and those of others showed some discrepancies in the expected densities. For example, the vapor density of sulphur was found to be about three times too great, that of phosphorus twice too great, that of mercury vapor and that of ammonium 303chloride only about half large enough to correspond to the values expected from analogy and other considerations. Thus, one volume of oxygen with two volumes of hydrogen make two volumes of steam, but only one third of a volume of sulphur vapor was found to unite with two volumes of hydrogen to make two volumes of hydrogen sulphide. Berzelius saw clearly that the results pointed to the existence of such molecules as S6, P4, and Hg1, but it was not generally realized in those days that Avogadro’s rule is fundamentally reliable, and Berzelius himself appears to have lost confidence in it on account of these complications, for he did not apply Avogadro’s principle to decisions about atomic weights, except in the cases of substances gaseous at ordinary temperatures.
Electro-chemical Theories.—The observation was made by Nicholson and Carlisle in 1800 that water was decomposed into its constituent gases by the electric current. Then in 1803 Berzelius and Hisinger found that salts were decomposed into their bases and acids by the same agency, and in 1807 Davy isolated potassium, sodium, and other metals afterwards, by a similar decomposition. Since those early times a vast amount of attention has been paid to the relation of electricity to chemical changes, a relation that is evidently of great importance from the fact that while electric currents decompose chemical compounds, these currents, on the other hand, are produced by chemical reactions.
Berzelius was particularly prominent in this direction, and in 1819 he published an elaborate electro-chemical theory. He believed that atoms were electrically polarized, and that this was the cause of their combination with one another. He extended this idea to groups of atoms, particularly to oxides, and regarded these groups as positive or negative, according to the excess of positive or negative electricity derived from their constituent atoms and remaining free. He thus arrived at his dualistic theory of chemical compounds, which attained great prominence and prevailed for a long time in chemical theory. According to this idea, each compound was supposed to be made up of a positive and a 304negative atom or group of atoms. For example, the formulas for potassium nitrate, calcium carbonate, and sulphuric acid corresponded to K2O.N2O5, CaO.CO2 and H2O.SO3 where we now write KNO3, CaCO3 and H2SO4, and the theory was extended to embrace organic compounds also.
The eminent English chemist and physicist Faraday announced the important law of electro-chemical equivalents in 1834. This law shows that the quantities of elements set free by the passage of a given quantity of electricity through their solutions correspond to the chemical equivalents of those elements. Faraday made a table of the equivalents of a number of elements, regarding them important in connection with atomic weights, but at that time no sharp distinction was usually made between equivalents and atomic weights, and it was not fully realized that one atom of a given element may be the electrical equivalent of several atoms of another.
Faraday’s law, which is still regarded as fundamentally exact, has been of much practical use in the measurement of electric currents and in calculations connected with electro-chemical processes. In discussing his experiments, Faraday made use of several new terms, such as “electrolyte” for a substance which conducts electricity when in solution, and is thus “electrolyzed,” “electrode,” “anode,” and “cathode,” terms that have come into general use, and finally “ions” for the particles that were supposed to “wander” towards the electrodes to be set free there.
This term “ion” remained in comparative obscurity for more than half a century, when it was brought into great prominence among chemists by Arrhenius in connection with the ionic theory.
Cannizzaro’s Ideas.—Up to about 1869 chaos reigned among the formulas used by different chemists. Various compound radicals and numerous type-formulas were employed, dualistic and unitary formulas of several kinds were in use, but the worst feature of the situation was the fact that more than one system of atomic weights was in vogue, so that water might be written
HO, H̶O, or H20
305and similar discrepancies might appear in nearly all formulas containing elements of different valencies. In 1858, however, an article by the Italian chemist Cannizzaro appeared in which the outlines of a course in chemical philosophy were presented. This acquired wide circulation in the form of a pamphlet at a chemical convention somewhat later, and it dealt so clearly and ably with Avogadro’s principle, Dulong and Petit’s law, and other points in connection with formulas that it led to a rapid and almost universal reform among those who were using unsatisfactory formulas.
At about this time also the dualistic formulas of Berzelius were generally abandoned, and hydrogen came to be regarded as the characteristic element of all acids. For instance, CaO.SO3, called “sulphate of lime,” came to be written CaSO4 and was called “calcium sulphate,” and while it had been shown as early as 1815 by Davy that “iodic acid,” I2O5, showed no acid reaction until it was combined with water, the accumulation of similar facts led to the formulation of sulphuric acid as H2SO4 instead of SO3 or H2O.SO3, and that of other “oxygen acids” in a similar way. As a necessary consequence of this view of acids, the bases came to be regarded as compounds of the “hydroxyl” group, OH. Therefore the formula for caustic soda came to be written NaOH instead of Na2O.H2O, and so on.
The Periodic System of the Elements.—The periodicity of the elements in connection with their atomic weights was roughly grasped by Newlands in England, who announced his “law of octaves” in 1863. This was at the time when the atomic weights were being modified and their numerical relations properly shown. The subject was worked out more fully by L. Meyer in Germany a little later, but it was most clearly and elaborately presented by the Russian chemist Mendeléeff in 1869.
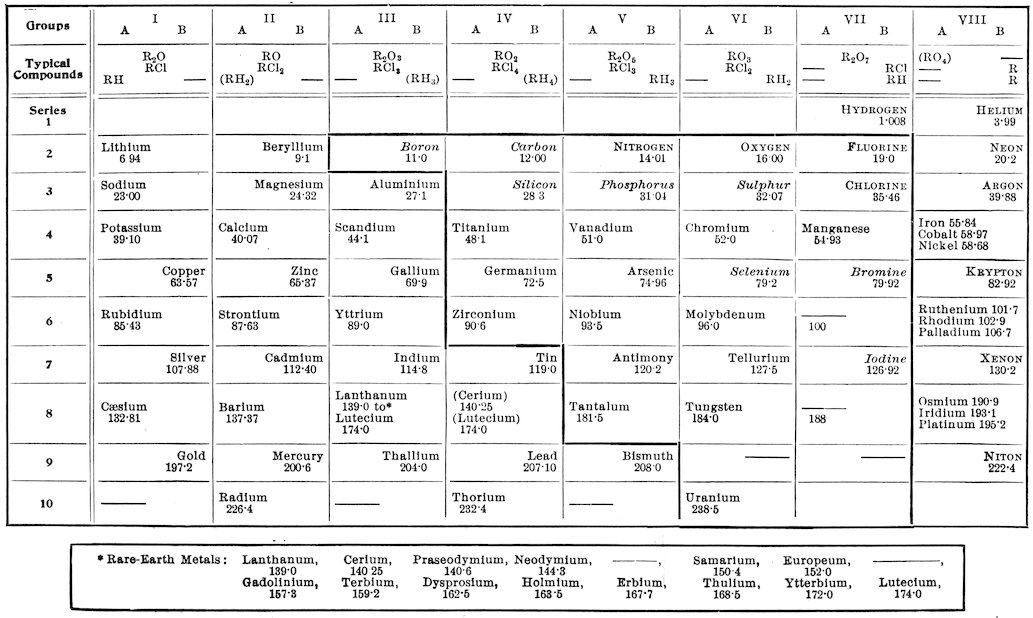
In order that this subject may be explained to some extent Mendeléeff’s table is given here, with the addition of the recently discovered elements and some other modifications.
306
Groups I II III IV V VI VII VIII
A B A B A B A B A B A B A B A B
Typical Compounds R2O RO R2O3 RO2 R2O5 RO3 R2O7 (RO4) —
RCl RCl2 RCl3 RCl4 RCl3 RCl2 — RCl — R
RH — (RH2) — — (RH3) — (RH4) — RH3 — RH2 — RH — R
Series 1 Hydrogen 1·008 Helium 3·99
2 Lithium 6·94 Beryllium 9·1 Boron 11·0 Carbon 12·00 Nitrogen 14·01 Oxygen 16·00 Fluorine 19·0 Neon 20·2
3 Sodium 23·00 Magnesium 24·32 Aluminium 27·1 Silicon 28·3 Phosphorus 31·04 Sulphur 32·07 Chlorine 35·46 Argon 39·88
4 Potassium 39·10 Calcium 40·07 Scandium 44·1 Titanium 48·1 Vanadium 51·0 Chromium 52·0 Manganese 54·93 Iron 55·84
Cobalt 58·97
Nickel 58·68
5 Copper 53·57 Zinc 65·37 Gallium 69·9 Germanium 72·5 Arsenic 74·96 Selenium 79·2 Bromine 79·92 Krypton 82·92
6 Rubidium 85·43 Strontium 87·63 Yttrium 89·0 Zirconium 90·6 Niobium 93·5 Molybdenum 96·0 —— 100 Ruthenium 101·7
Rhodium 102·9
Palladium 106·7
7 Silver 107·88 Cadmium 112·40 Indium 114·8 Tin 119·0 Antimony 120·2 Tellurium 127·5 Iodine 126·92 Xenon 130·2
8 Cæsium 132·81 Barium 137·87 Lanthanum 139·0 to* Lutecium 174.0 (Cerium) 140·25 (Lutecium) 174.0 Tantalum 181·5 Tungsten 184·0 —— 188 Osmium 190·9
Iridium 193·1
Platinum 195·2
9 Gold 197·2 Mercury 200·6 Thallium 204·0 Lead 207·10 Bismuth 208·0 Niton 222·4
10 Radium 226·4 Thorium 292·4 Uranium 238·5
* Rare·Earth Metals: Lanthanum, 139·0 Cerium, 140·25 Praseodymium, 140·6 Neodymium, 144·3 ——, Samarium, 150·4 Europeum, 152·0 ——,
Gadolinium, 157·3 Terbium, 159·2 Dysprosium, 162·5 Holmium, 163·5 Erbium, 167·7 Thulium, 168·5 Ytterbium, 172·0 Lutecium, 174·0
Note.—Distinctions in printing: Gaseous elements. Other non-metallic elements, metallic elements. The heavy line encloses approximately the acid-forming elements.
307In this table the elements arranged in the order of their atomic weights fall into eight groups where the known oxides progress regularly, with the exception of two or three elements, from R2O in Group I to R2O7 in Group VII, while in Group VIII two oxides (of ruthenium and osmium) are known which carry the progression to RO4.
It was pointed out by Mendeléeff that, with the exception of series 1 and 2 at the top of the table, the alternate members of the groups show particularly close relationships. These subordinate groups, marked A and B, in most cases show remarkable analogies and gradations in their properties, for example, in the alkali-metals from lithium to cæsium, and in the halogens from fluorine to iodine. The two divisions of a group do not usually show very close relations to each other, except in their valency, and they even display, in several instances, opposite gradations in chemical activity in the order of their atomic weights. For instance, cæsium stands at the electro-positive end, while gold stands at the electro-negative end of its subordinate group. The difference between the two divisions is very great in Groups VI and VII, but it is extreme in Group VIII, where heavy metals are on one side and inactive gases on the other. Many authorities separate these gases into a “Group O” by themselves at the left-hand side of the table, but this does not change their relative positions, and the plan may be objected to on the ground that many vacant places are thus left in the groups VIII and O.
The periodic law has been useful in rectifying certain atomic weights. At the outset Mendeléeff was obliged to change beryllium from 14·5 (assuming Be2O3) to 9 (assuming BeO), and later the atomic weights of indium and uranium were changed to make them fit the system. All of these changes have been confirmed by physical means.
Mendeléeff found a number of vacant places in his table, and was thus able to render further service to chemical science by predicting the properties of undiscovered elements, and his predictions were very closely confirmed by the later discovery of scandium, gallium, and germanium. The table indicates that there are still 308two undiscovered elements below manganese and probably two more among the rare-earth metals. The interesting observation has just recently been made by Soddy that the products of radioactive disintegration appear to pass in a symmetrical way through positions in the periodic system, giving off a helium molecule at alternate transformations until the place of lead is reached. It appears, therefore, that the five vacant places in the table above bismuth are probably occupied by these evanescent elements, and it is to be noticed that all of the elements that have been placed in this region of high atomic weights are radioactive.
There are some inconsistencies in the periodic system. The increments in the atomic weights are irregular, and there are three cases, argon and potassium, cobalt and nickel, and tellurium and iodine, where a higher atomic weight is placed before a lower one in order to bring these elements into their undoubtedly proper places. There is a peculiarity also in the heavy-metal division of Group VIII, where three similar elements occur in each of three places, and where the usual periodicity appears to be suspended, or nearly so, in comparison with most of the other elements. However, there seems to be a still more remarkable case of this kind in Group III, where fourteen metals of the rare earths have been placed. They are astonishingly similar in their chemical properties, hence it seems necessary to assume that periodicity is suspended here throughout the wide range of atomic weights from 139 to 174, where no elements save these have been found.
Several other interesting features of the table may be pointed out. The chlorides and hydrides, as indicated by the “typical compounds,” show a regular progression in both directions towards Group IV. (Where the type-formulas do not apply, as far as is known, to more than one or two elements, they have been placed in parentheses in the table given here.) It is a striking fact that the acid-forming elements occur together in a definite part of the table, and that the gases and other non-metallic elements, except the inactive gases of Group VIII, occur in the same region.
309Atomic Numbers.—As the result of a spectroscopic study of the wave lengths or frequencies of the X-rays produced when cathode rays strike upon anticathodes composed of different elements, Moseley in 1914 discovered that whole numbers in a simple series can be attributed to the atoms. These atomic numbers are: 1 for hydrogen, 2 for helium, 3 for lithium, 4 for beryllium, and so on, in the order in which the elements occur in Mendeléeff’s periodic table, and in the cases of argon and potassium, cobalt and nickel, and tellurium and iodine, they follow the correct chemical order, while the atomic weights do not. They appear to indicate, therefore, an even more fundamental relation between the atoms than that shown by the atomic weights.
These numbers are now available for every element up to lead, and they are particularly interesting in indicating, on account of missing numbers, the existence of two undiscovered elements in the manganese group, and two more among the rare-earth metals, in confirmation of the vacant places below lead in Mendeléeff’s table.
The Isolation of Elements.—In the year 1818 about 53 elements were recognized, and since that time about 30 more have been discovered, but the elements already known comprised the more common ones, and nearly all of those which have been commercially important. A few of them, including beryllium, aluminium, silicon, magnesium, and fluorine, were then known only in their compounds, as they had not yet been isolated in the free condition.
Berzelius in 1823 prepared silicon, a non-metallic element resembling carbon in many respects. This element has recently been prepared on a rather large scale in electric furnaces at Niagara Falls, and has been used for certain purposes in the form of castings.
Wöhler created much sensation in 1827 by isolating aluminium and finding it to be a very light, strong and malleable metal, stable in the air, and of a silver-white color. For a long time this metal was a comparative rarity, being prepared by the reduction of aluminium chloride with metallic sodium; but about 25 years ago Hall, an American, devised a method of preparing it by 310electrolyzing aluminium oxide dissolved in fused cryolite. This process reduced the cost of aluminium to such an extent that it has now come into common use.
Wöhler and Bussy prepared beryllium in 1828, and Liebig and Bussy did the same service for magnesium in 1830. The latter metal has come to be of much practical importance, both as a very powerful reducing agent in chemical operations, and as an ingredient of flash-light powders and of mixtures used for fireworks. It is also used in making certain light alloys.
After almost innumerable attempts to isolate fluorine, during a period of nearly a century, this was finally accomplished in 1886 by Moissan in France by the electrolysis of anhydrous hydrogen fluoride. The free fluorine proved to be a gas of extraordinary chemical activity, decomposing water at once with the formation of hydrogen fluoride and ozonized oxygen. This fact explains the failure of many previous attempts to prepare it in the presence of water.
Early Discoveries of New Elements.—The remarkable activity of chemical research at the beginning of our period is illustrated by the fact that three new elements were discovered in 1817. In that year Berzelius had discovered selenium, Arfvedson, working in Berzelius’s laboratory had discovered the important alkali-metal lithium, and Stromeyer had discovered cadmium.
In 1826 Ballard in France discovered bromine in the mother-liquor from the crystallization of common salt from sea water. Bromine proved to be an unusually interesting element, being the only non-metallic one that is liquid at ordinary temperatures, and being strikingly intermediate in its properties between chlorine and iodine. It has been obtained in large quantities from brines, and is produced extensively in the United States. The elementary substance and its compounds have found important applications in chemical operations, while the bromides have been found valuable in medicine and silver bromide is very extensively used in photography.
In 1828 Berzelius discovered thorium. The oxide of this metal has recently been employed extensively as the principal constituent of incandescent gas-mantles, and the element has acquired particular importance from the 311fact that, like uranium, it is radioactive, decomposing spontaneously into other elements.
Vanadium had been encountered as early as 1801 by Del Rio, who named it “erythronium,” but a little later it was thought to be identical with chromium and was lost sight of for a while. In 1830, however, it was re-discovered by, and received its present name from Sefström in Sweden. Berzelius immediately made an extensive study of vanadium compounds, but he gave them incorrect formulas and derived an incorrect atomic weight for the element, because he mistook a lower oxide for the element itself. Roscoe in England in 1867 isolated vanadium for the first time, found the right atomic weight, and gave correct formulas to its compounds. Vanadium is particularly interesting from the fact that it displays several valencies in its compounds, many of which are highly colored. It has found important use as an ingredient in very small proportions in certain “special steels” to which it imparts a high degree of resistance to rupture by repeated shocks.
Columbium was discovered early in the nineteenth century in the mineral columbite from Connecticut by Hatchett, an Englishman, who did not, however, obtain the pure oxide. It was afterwards obtained by Rose who named it niobium. Both names for the element are in use, but the former has priority. Attention was called to this fact by an article in the Journal by Connell, an Englishman (18, 392, 1854).
The Platinum Group of Metals.—In 1854 a new member of the platinum group of metals, ruthenium, was discovered by Claus. Platinum had been discovered about the middle of the eighteenth century, while its other rarer associates, iridium, osmium, palladium, and rhodium, had been recognized in the very early years of the nineteenth century. It was during the latter period that platinum ware began to be employed to a considerable extent in chemical operations, and this use was greatly extended as time went on. The discovery was made by Phillips in 1831 that finely divided platinum by contact would bring about the combination of sulphur dioxide with atmospheric oxygen, and this application during the past 20 years has become enormously important in the sulphuric 312acid industry, while other important applications of platinum as a “catalytic agent” have also been made. Wolcott Gibbs and Carey Lea have contributed perhaps more than any other recent chemists to a knowledge of the platinum metals. Carey Lea (38, 81, 248, 1864) dealt chiefly with the separation of the metals from each other, while Gibbs’s work (31, 63, 1861; 34, 341, 1862) included investigations of many of the compounds.
It may be mentioned that while platinum and its associates were formerly known only in the uncombined condition in nature, the arsenide sperrylite, PtAs2, was described by the late S. L. Penfield, and the senior writer of this chapter, in articles published in the Journal (37, 67, 71, 1889).
Applications of the Spectroscope.—The discovery in certain mineral waters of the rare alkali-metals rubidium and cæsium by Bunsen and Kirchoff in 1861 was in consequence of the application of spectroscopy by these same scientists a short time previously to the identification of elements imparting colors to the flame. Since that time the employment of the spectroscope for chemical purposes has been much extended, as it has been used in the examination of light from electric sparks and arcs, as well as from Geissler tube discharges and from colored solutions.
The metals rubidium and cæsium are interesting in being closely analogous to potassium and in standing at the extreme electro-positive end of the series of known metals. It should be noticed here that Johnson and Allen of our Sheffield Laboratory, having obtained a good supply of rubidium and cæsium material from the lepidolite of Hebron, Maine, made some important researches upon these elements, accounts of which were published in the Journal (34, 367, 1862; 35, 94, 1863). They established the atomic weight of cæsium, thus correcting Bunsen’s determination which was unsatisfactory on account of the small quantity and impurity of his material. Pollucite, a mineral rich in cæsium, which had been found in very small amount on the Island of Elba, has more recently been obtained in large quantities—hundreds of pounds—at Paris, Maine, and its vicinity. This American pollucite was first analyzed and identified 313by the senior writer of this article (41, 213, 1891), and later (43, 17, 1892 et seq.) the results of many investigations on cæsium and rubidium compounds, in which the junior writer played an important part, carried out in Sheffield Laboratory, were published in the Journal.
The application of the spectroscope led to the discovery of thallium in 1861 by Crookes of England, and to that of indium in 1863 by Reich and Richter in Germany. Both of these metals are extremely rare, but they are of considerable theoretical interest. Thallium is particularly remarkable in showing resemblances in its different compounds to several groups of metals.
The spectroscope was employed again in connection with the discovery of gallium in 1875 by Boisbaudran. It is in the same periodic group as thallium and indium, and it has a remarkably low melting point, just above ordinary room-temperature. It has been among the rarest of the rare elements, but within two or three years a source of it has been found in the United States in certain residues from the refining of commercial zinc. The recent issues of the Journal (41, 351, 1916; 42, 389, 1916) show that Browning and Uhler of Yale have availed themselves of this new material in order to make important chemical and physical researches upon this metal.
Germanium.—The discovery of germanium in the mineral argyrodite in 1886 by Winkler revealed a curious metal which gives a white sulphide that may be easily mistaken for sulphur and which is volatilized completely when its hydrochloric acid solution is evaporated, so that it is evasive in analytical operations. This element had been predicted with much accuracy by Mendeléeff, and it is rather closely related to tin.
A few years after the discovery of germanium, Penfield published in the Journal (46, 107, 1893; 47, 451, 1894) some analyses of argyrodite, correcting the formula given by Winkler to the mineral; also he described canfieldite, an analogous mineral from Bolivia, in which a large part of the germanium was replaced by tin.
The Rare Earths.—Before the year 1818 two rare earths, the oxides of yttrium and cerium, were known in an impure condition. Since that time about fourteen others have been discovered as associates of the first 314two. The rare earths are peculiar from the fact that many of them are always found mixed together in the minerals containing them, and also from the circumstance that most of them are remarkably similar in their chemical reactions and consequently exceedingly difficult to separate from each other. In many cases multitudes of fractional precipitations or crystallizations are needed to obtain pure salts of a number of these metals. The solutions of the salts of several of these elements give characteristic absorption bands when examined spectroscopically by the use of transmitted light.
No important practical application has been found for any of these earthy oxides, except that about one per cent of cerium oxide is mixed with thorium oxide in incandescent gas-mantles in order to obtain greatly increased luminosity.
The Inactive Gases.—As long ago as 1785, Cavendish, that remarkable Englishman who first weighed the world and first discovered the composition of water, actually obtained a little argon in a pure condition by sparking atmospheric nitrogen with oxygen converting it into nitric acid (another discovery of his) and absorbing the excess of oxygen. The volume of this residual gas as estimated by him corresponds very closely to the volume of argon in the atmosphere, as now known.
It was more than a century later, in 1894, that Rayleigh and Ramsay discovered argon in the air. Lord Rayleigh had found that atmospheric nitrogen was about one-half per cent heavier than chemical nitrogen, a fact which led to the investigation. It was only necessary to repeat Cavendish’s experiment on a large scale, or to absorb oxygen with hot copper and nitrogen with hot magnesium, in order to obtain argon. The gas attracted much attention, both on account of having but a single atom in its molecule, and particularly because it failed to enter into chemical combination of any kind. This gas has been used of late for filling the bulbs of incandescent electric lamps in cases where a gas pressure without chemical action is desired.
In 1890 and 1891, Hillebrand published in the Journal (40, 384, 1890: 42, 390, 1891) a series of analyses of the mineral uraninite and reported in some samples of the 315mineral as much as 2·5 per cent of an inactive gas. Hillebrand examined the gas spectroscopically but, just missing an important discovery, he detected only the spectrum lines of nitrogen. Ramsay, in searching for argon in some sort of natural combination, and doubtless remembering Hillebrand’s work, heated some cleveite, a variety of uraninite, and obtained, not argon, but a new gas. This gave a yellow spectrum-line corresponding to a line previously observed in the light of the sun’s corona and attributed to an element in the sun called helium. Helium, therefore, in 1895 had been found on the earth. This gas is a constant constituent of uranium minerals, as it is produced by the breaking down of radioactive elements. It has been found in very small quantity in the atmosphere, and is the most difficult of all known gases to liquefy, as its boiling point, as shown by Onnes in 1908, is only 4° above the absolute zero. It has not yet been solidified.
In 1898 Ramsay and Travers, by the use of ingenious methods of fractional distillation and absorption by charcoal, obtained three other much rarer inactive gases from the atmosphere which they called neon, krypton and xenon.
The inactive gases are all colorless, and as they form no chemical compounds they are characterized by their densities, which give their atomic weights, by their boiling points, and by their characteristic Geissler-tube spectra.
The gaseous radium emanation, or niton, belongs also to the inactive group, and it was also collected and studied by Ramsay who was compelled to work with only 0·0001 cc. of it, as the volume obtained by heating radium salts is very small. It is an evanescent element, disappearing within a few days on account of radioactive disintegration. Meanwhile it glows brilliantly when liquefied and cooled to the temperature of liquid air. It has an atomic weight of 222, four units below that of radium, and the difference is considered as due to the loss by radium of an atom of helium in passing into the emanation.
The Radioactive Elements.—The discovery of radium in 1898 by Madame Curie, and the study of that and other 316radioactive elements has produced a profound effect upon chemical theory. It was found that the two elements of the highest atomic weights, uranium and thorium, are always spontaneously decomposing into other elements at a fixed rate of speed which can be controlled by no artificial means, and that the elements resulting from these decompositions likewise undergo spontaneous changes into still other elements at greatly varying rates of speed, forming in each case a remarkable series of temporary elements. These transformations are accompanied by the emission at enormous velocities of three kinds of rays, one variety of which has been shown to consist of helium atoms. The greater number of the elements formed in these transformations have not as yet been obtained in a pure condition, and they are known only in connection with their radioactivity, volatility, etc.; but radium and niton, two of these products, have been obtained in a pure condition, so that their atomic weights and their places in the periodic system have been fixed.
We owe much of our knowledge of the radioactive transformations to the researches of Rutherford and of Soddy, and of their co-workers, but one of the important products of the transformation of uranium, an element which he called ionium, was characterized by Boltwood of Yale (25, 365, 1908).
Radium and niton, apart from their radioactive properties, resemble barium and the inert gases of the atmosphere, respectively. The rates at which their progenitors produce them, and the rates at which they themselves decompose, bring about a state of equilibrium after a time. Therefore a given amount of uranium, which decomposes exceedingly slowly, can yield even after thousands of years only a very small proportional quantity of undecomposed radium, one-half of which disappears in about 2500 years, because the amount decomposed must eventually be equal to the amount produced. The first conclusive evidence that radium is a product of the decomposition of uranium was given by Boltwood in the Journal (18, 97, 1904). He found that all uranium minerals contain radium; and the amount of radium present is always proportional to the amount 317of uranium, which shows the genetic relation between the two.
In the case of niton, which is produced by radium, and is called also the radium emanation, the rate of decay is rapid, so that if the gas is expelled from radium by heating, equilibrium is reached after a few days, with the accumulation of the largest possible amount of niton.
The conclusion has been reached by Rutherford and others that the final product besides helium, in the radioactive transformations, is lead, or at least an element or elements resembling lead to such a degree that no separation of them by chemical means is possible. Atomic weight determinations by Richards and others have shown that specimens of lead found in radioactive minerals give distinctly different atomic weights from that of ordinary lead. This fact has led to the view that possibly the atoms of the elements are not all of the same weight, but vary within certain limits—a view that is contrary to previous conclusions derived from the uniformity in atomic weights obtained with material from many different sources.
The results of the investigations upon radioactivity have led to modified views in regard to the stability of the elements in general. There has been little or no proof obtained that any artificial transmutation of the elements is possible, but the spontaneous transformation of the radioactive elements brings forward the possibility that other elements are changing imperceptibly, and that a state of evolution exists among them. All of the radioactive changes that we know proceed from higher to lower atomic weights, and we are entirely ignorant of the process by which uranium and thorium must have been produced originally.
Since radioactive changes have been found to be accompanied by the release of vast amounts of energy, compared with which the energy of chemical reactions is trivial, a new aspect in regard to the structure of atoms has arisen,—they must be complex in structure, the seats of enormous energy.
The determination of the amount of radium in the earth’s crust has indicated that the heat produced by it is amply sufficient to supply the loss of heat due to radiation, 318and this source of heat is regarded by many as the cause of volcanic action. The sun’s radiant heat also has been supposed to be supplied by radioactive action, so that the older views regarding the limitation of the age of the earth and the solar system on account of loss of heat have been considerably modified by our knowledge of radioactivity.
Physical Chemistry.
The application of physical methods as aids to chemical science began in early times, and some of these, such as the determinations of gas and vapor densities, specific heats, and crystalline forms have been mentioned already in this article. Within recent times physical chemistry has greatly developed and a few of its important achievements will now be described.
Molecular Weight Determinations.—Gas and vapor densities in connection with Avogadro’s principle, formed the only basis for molecular weight determinations until comparatively recent times. The early methods of Gay-Lussac and Dumas for vapor density were supplemented in 1868 by the method of Hofmann, whereby vapors were measured under diminished pressure over mercury. In 1878 Victor Meyer introduced a simpler method depending upon the displacement of air or other gas by the vapor in a heated tube. As refractory tubes, such as those of porcelain or even iridium, could be used in this method, molecular weights at extremely high temperatures were determined with interesting results. For instance, it was found that iodine vapor, which shows the molecule I2 at lower temperatures, gradually becomes monatomic with rise in temperature, that sulphur vapor dissociates from S8 to S2 under similar conditions, and that most of the metals, including silver, have monatomic vapors.
In 1883 and later it was pointed out by Raoult that the molecular weights of substances could be found from the freezing points of their solutions, but this method was complicated from the fact that salts, strong acids and strong bases behaved quite differently from other substances in this respect, and allowances had to be made for the types of substances used. The complication was 319afterwards explained by the ionization theory of Arrhenius. Better apparatus for this method was soon devised by Beckmann, who introduced also a method depending upon the boiling points of solutions, and these two methods are still the standard ones for determining molecular weights in solution. They are very extensively employed by organic chemists.
It has been found that the majority of substances when dissolved have the same molecular weight as in the gaseous condition, provided that they can be volatilized at comparable temperatures. For instance, sulphur in solution has the formula S8, iodine is I2 and the metals are monatomic.
Van’t Hoff’s Law and Arrhenius’s Theory of Ions.—Modern views on solutions date largely from 1886, when van’t Hoff called attention to the relations existing between the osmotic pressure exerted by dissolved substances and gas pressure.
Pfeffer, a botanist, was the first to measure osmotic pressure (1877). Basing his conclusions chiefly upon Pfeffer’s determinations, van’t Hoff formulated a new and highly important law, which may be stated as follows: The osmotic pressure exerted by a substance in solution is equal to the gas pressure that the substance would exert if it were a gas at the same temperature and the same volume. Further investigations have fully established the fact that molecules in dilute solution obey the simple laws of gases.
It was pointed out by van’t Hoff that salts, strong acids and strong bases showed marked exceptions to his law in exerting much greater osmotic pressures than those calculated for them.
The next year in 1887, Arrhenius explained this abnormal behavior of salts, strong acids and strong bases by assuming that they dissociate spontaneously into ions when they dissolve, and that these more numerous particles act like molecules in producing osmotic pressure. He showed that these exceptional substances all conduct electricity in solution, while those conforming with van’t Hoff’s law do not, and according to his theory the ions become positively or negatively charged when they are formed, and these charged ions conduct the current. 320For example a molecule of sodium chloride was supposed to give the two ions Na+ and Cl-, thus exerting twice as much osmotic pressure as a single molecule.
Determinations of osmotic pressure or related values, such as depression of the freezing point and of electric conductivity, indicated that ionization could not be regarded as complete in any case except in exceedingly dilute solutions, and that the extent of ionization varied with different substances. The fact that osmotic pressures and electric conductivities gave closely agreeing results in regard to the extent of ionization in various cases, is the strongest evidence in support of the theory.
It was difficult at first for many chemists to believe that atoms, such as those of sodium and chlorine, and groups such as NH4 and SO4 could exist independently in solution, even though electrically charged. However, the theory rapidly gained ground and is now accepted by nearly every chemist as a satisfactory explanation of many facts.
During recent years, many investigations relating to osmotic pressure and ionization have been carried out in the United States, but only the work of Morse, A. A. Noyes, and the late H. C. Jones can be merely alluded to here. It should be mentioned that the eminent author of the ionic hypothesis gave the Silliman Memorial course of lectures at Yale in 1911 on Theories of Solution.
Colloidal Solutions.—Graham, an English chemist, in 1861 was the first to make a distinction between substances forming true solutions, which he called crystalloids, and those of a gummy nature resembling glue, which in solution do not diffuse readily through parchment membranes, as crystalloids do, and which he called colloids. The separation of colloids by means of parchment was called dialysis, and this process has come into extensive use in preparing pure colloidal solutions. Slow diffusion is now regarded as characteristic of colloids rather than their gummy condition.
Colloidal solutions occupy an intermediate position between true solutions and suspensions, resembling one or the other according to the kind of colloid and the fineness of division. By preparing filters with pores of varying degrees of fineness, Bechold has been able to 321separate colloids from each other in accordance with the size of their particles. It has also been possible to prepare different solutions of a colloid varying gradually from one in which the particles were undoubtedly in suspension to one which had many of the properties of a true solution.
Beginning in 1889, Carey Lea described in the Journal (37, 476, 1889 et seq.) a variety of methods for preparing colloidal solutions of the metals, consisting in general of treating solutions of metallic salts with mild reducing agents. His work on colloidal silver was particularly extensive and interesting. Solutions of this kind have recently yielded some extremely interesting results by means of the ultra-microscope, an apparatus devised by Zsigmondy and Siedentopf. A very intense beam of light is passed through the solution and observed at right angles with a powerful microscope. Under these conditions, particles much too small to be seen by other means, reveal their presence by reflected light. It has been possible in a very dilute solution of known strength to count the particles and thus to calculate their size. The smallest colloidal particles measured in this way were of gold and were shown to have approximately ten times the diameter, or 1000 times the volume, attributed to ordinary molecules. It is of interest that the particles appear in rapid motion corresponding to the well-known Brownian movement.
The chemistry of colloids has now assumed such importance that it may be considered as a separate branch of the science. It has its own technical journal and deals largely with the chemistry of organic products. All living matter is built up of colloids, and hæmoglobin, starch, proteins, rubber and milk are examples of colloidal substances or solutions. Among inorganic substances, many sulphides, silicic acid, and the amorphous hydroxides, like ferric hydroxide, frequently act as colloids.
Law of Mass Action.—Berthollet about the beginning of the last century was the first chemist to study the effect of mass, or more correctly, the concentration of substances on chemical action. His views summarized by himself are as follows: “The chemical activity of a 322substance depends upon the force of its affinity and upon the mass which is present in a given volume.” The development of this idea, which is fundamentally correct, was greatly hindered by the fact that Berthollet drew the incorrect conclusion that the composition of chemical compounds depended upon the masses of the substances combining to produce them, a conclusion in direct contradiction to the law of definite proportions, and since this view was soon disproved by Proust and others, Berthollet’s law in its other applications received no immediate attention. Mitchell, however, pointed out in the Journal (16, 234, 1829) the importance of Berthollet’s work, and Heinrich Rose in 1842 again called attention to the effect of mass, mentioning as one illustration the effect of water and carbonic acid in decomposing the very stable natural silicates. Somewhat later several other chemists made important contributions to the question of the influence of concentration upon chemical action, but it was the Norwegians, Guldberg and Waage, who first formulated the law of mass action in 1867.
This law has been of enormous importance in chemical theory, since it explains a great many facts upon a mathematical basis. It applies particularly to equilibrium in reversible reactions, where it states that the product of the concentrations on the one side of a simple reversible equation bears a constant relation to the products of the concentrations on the other side, provided that the temperature remains constant. In cases of this kind where two gases or vapors react with two solids, the latter if always in excess may be regarded as constant in concentration, and the law takes on a simpler aspect in applying only to the concentrations of the gaseous substances. For example, in the reversible reaction
3Fe + 4H2O ⇄ Fe3O4 + 4H2,
which takes place at rather high temperatures, a definite mixture of steam and hydrogen at a definite temperature will cause the reaction to proceed with equal rapidity in both directions, thus maintaining a state of equilibrium, provided that both iron and the oxide are present in 323excess. If, however, the relative concentrations of the hydrogen and steam are changed, or even if the temperature is changed, the reaction will proceed faster in one direction than in the other until equilibrium is again attained.
The principle of mass action also explains why it is sometimes possible for a reversible reaction to become complete in either direction. For instance, in connection with the reaction that has just been considered, if steam is passed over heated iron and if hydrogen is passed over the heated oxide, the gaseous product in each case is gradually carried away, and the reaction continually proceeds faster in one direction than in the other until it is complete, according to the equations
3Fe + 4H2O → 3Fe3O4 + 4H2, and
Fe3O4 + 4H2 → 3Fe + 4H2O.
Many other well-known and important facts, both chemical and physical, depend upon this law. It explains the circumstance that a vapor-pressure is not dependent upon the amount of the liquid that is present; it also explains the constant dissociation pressure of calcium carbonate at a given temperature, irrespective of the amounts of carbonate and oxide present; in connection with the ionic theory, it furnishes the reason for the variable solubility of salts due to the presence of electrolytes containing ions in common; and it elucidates Henry’s law which states that the solubilities of gases are proportional to their pressures.
Ostwald, more than any other chemist, has been instrumental in making general applications of this law, and he made particularly extensive use of it in connection with analytical chemistry in a book upon this subject which he published.
The Phase Rule.—In 1876 Willard Gibbs of Yale published a paper in the Proceedings of the Connecticut Academy of Science on the “Equilibrium of Heterogeneous Substances,” and two years later he published an abstract of the article in the Journal (16, 441, 1878). He had discovered a new law of nature of momentous importance and wide application which is called the 324“Phase-Rule” and is expressed by a very simple formula.
The application of this great discovery to chemical theory was delayed for ten years, partly, perhaps, because it was not sufficiently brought to the attention of chemists, but largely it appears because it was not at first understood, since its presentation was entirely mathematical.
It was Rooseboom, a Dutch chemist, who first applied the phase-rule. It soon attracted profound attention, and the name of Willard Gibbs attained world-wide fame among chemists. When Nernst, who is perhaps the most eminent physical chemist of the present time, was delivering the Silliman Memorial Lectures at Yale a few years ago, he took occasion to place a wreath on the grave of Willard Gibbs in recognition of his achievements.
To understand the rule, it is necessary to define the three terms, introduced by Gibbs, phase, degrees of freedom and component.
By the first term, is meant the parts of any system of substances which are mechanically separable. For instance, water in contact with its vapor has two phases, while a solution of salt and water is composed of but one. The degrees of freedom are the number of physical conditions, including pressure, temperature and concentration, which can be varied independently in a system without destroying a phase. The exact definition of a component is not so simple, but in general, the components of a system are the integral parts of which it is composed. Any system made up of the compound H2O, for instance, whether as ice, water or vapor, contains but one component, while a solution of salt and water contains two. Letting P, F, and C stand for the three terms, the phase-rule is simply
F = C + 2 − P
that is, the number of degrees of freedom in a system in equilibrium equals the number of components, plus two, minus the number of phases. The rule can be easily understood by means of a simple illustration. In a system composed of ice, water and water vapor, there are three phases and one component and therefore
F = 1 + 2 − 3 = 0
Such a system has no degrees of freedom. This means that no physical condition, pressure or temperature can be varied without destroying a phase, so that such a system can only exist in equilibrium at one fixed temperature, with a fixed value for its vapor-pressure.
325For instance, if the system is heated above the fixed temperature, ice disappears and if the pressure is raised, vapor is condensed. If this same system of water alone contains but two phases, for instance, liquid and vapor, F = 1 + 2 − 2 = 1, or there is one degree of freedom. In such a system, one physical condition such as temperature can be varied independently, but only one, without destroying a phase. For instance, the temperature may be raised or lowered, but for every value of temperature there is a corresponding value for the vapor-pressure. One is a function of the other. If both values are varied independently, one phase will disappear, either vapor condensing entirely to water or the reverse. Finally if the system consists of one phase only, as water vapor, F = 2, or the system is divariant, which means that at any given temperature it is possible for vapor to exist at varying pressures.
The illustration which has been given relates to physical equilibrium, but the rule is applicable to cases involving chemical changes as well. In comparing the phase-rule with the law of mass action, it will be noticed that both have to do with equilibrium. The great advantage of the former is that it is entirely independent of the molecular condition of the substances in the different phases. For instance, it makes no difference so far as the application of the rule is concerned, whether a substance in solution is dissociated, undissociated or combined with the solvent. In any case, the solution constitutes one phase. On the other hand, the rule is purely qualitative, giving information only as to whether a given change in conditions is possible. The law of mass action is a quantitative expression so that when the value of the constant is once known, the change can be calculated which takes place in the entire system if the concentration of one substance is varied. The law, however, requires a knowledge of the molecular condition of 326the reacting substances, which may be uncertain or unknown, and chiefly on this account it has, like the phase-rule, often only a qualitative significance.
The phase rule has served as a most valuable means of classifying systems in equilibrium and as a guide in determining the possible conditions under which such systems can exist. As illustrations of its practical application, van’t Hoff used it as an underlying principle in his investigations on the conditions under which salt deposits have been formed in nature, and Rooseboom was able by its means to explain the very complicated relations existing in the alloys of iron and carbon which form the various grades of wrought iron, steel and cast iron.
Thermochemistry.—This branch of chemistry has to do with heat evolved or absorbed in chemical reactions. It is important chiefly because in many cases it furnishes the only measure we have of the energy changes involved in reactions. To a great extent, it dates from the discovery by Hess in 1840 of a fundamental law which states that the heat evolved in a reaction is the same whether it takes place in one or in several stages. This law has made it possible to calculate the heat values of a large number of reactions which cannot be determined by direct experiment.
Thermochemistry has been developed by a comparatively few men who have contributed a surprisingly large number of results. Favre and Silbermann, beginning shortly after 1850, improved the apparatus for calorimetric determinations, which is called the calorimeter, and published many results. At about the same time Julius Thomsen, and in 1873 Berthelot, began their remarkable series of publications which continued until recently. Thomsen’s investigations were published in 1882 in 4 volumes. It is probably safe to say that the greater part of the data of thermochemistry was obtained by these two investigators. The bomb calorimeter, an apparatus for determining heat values by direct combustion, was developed by Berthelot. The recent work of Mixter at Yale, published in the Journal, and of Richards at Harvard should be mentioned particularly. Mixter’s work in this field began in 1901 (12, 347). Using an improved bomb calorimeter, he has developed a 327method of determining the heats of formation of oxides by combustion with sodium peroxide. By this same method as well as by direct combustion in oxygen, he has obtained results which appear to equal or excel in accuracy any which have ever been obtained in his field of work. Richards’s work has consisted largely of improvements in apparatus. He developed the so-called adiabatic calorimeter which practically eliminates one of the chief errors in thermal work caused by the heating or cooling effect of the surroundings. This modification is being generally adopted where extremely accurate work is required.
Organic Chemistry.
One hundred years ago qualitative tests for a few organic compounds were known, the elements usually occurring in them were recognized, and some of them had been analyzed quantitatively, but organic chemistry was far less advanced than inorganic, and almost the whole of its enormous development has taken place during our period.
Berzelius made a great advance in the subject by establishing the fact, which had been doubted previously, that the elements in organic compounds are combined in constant, definite proportions. In 1823 Liebig brought to light the exceedingly important fact of isomerism by showing that silver fulminate had the same percentage composition as silver cyanate, a compound of very different properties. Isomeric compounds with identical molecular weight as well as the same composition have since been found in very many cases, and they have played a most important part in determining the arrangements of atoms in molecules. They have been found to be very numerous in many cases. For instance, three pentanes with the formula C5H12are known, all that are possible according to theory, and in each case the structure of the molecule has been established. On theoretical grounds it has been calculated that 802 isomeric compounds with the formula C13H28 are possible, while with more complex formulas the numbers of isomers may be very much greater.
328A particularly interesting case of isomerism was observed by Wöhler in 1828, when he found that ammonium cyanate changes spontaneously into urea
(NH4CNO → N2H4CO).
This was the first synthesis of an organic compound from inorganic material, and it overthrew the prevailing view that vital forces were essential in the formation of organic substances. A great many natural organic compounds have been made artificially since that time, and some of them, such as artificial alizarin, indigo, oil of wintergreen, and vanillin, have more or less fully replaced the natural products. The preparation of a vast number of compounds not known in nature, many of which are of practical importance as medicines, dyes, explosives, etc., has been another great achievement of organic chemistry.
The development of our present formulas for organic compounds, by means of which in many cases the relative positions of the atoms can be shown with the greatest confidence, has been gradual. Formulas based on the dualistic idea of Berzelius were used for some time, type-formulas, with the employment of compound radicals, came later, the substitution of atoms or groups of atoms for others in chemical reactions came to be recognized, but one of the most important steps was the recognition of the quadrivalence of carbon and the general application of valency to atoms by Kekulé about 1858. This led directly to the use of modern structural formulas which have been of the greatest value in the theoretical interpretation of organic reactions. It was Kekulé also who proposed the hexagonal ring-formula for benzene, C6H6, which led to exceedingly important theoretical and practical developments. The details of the formulas for many other rings and complex structures have been established since that time, and there is no doubt that the remarkable achievements in organic chemistry during the past sixty years have been much facilitated by the use of these formulas.
Many important researches in organic chemistry have been carried out in the United States, and the activity in this direction has greatly increased in recent years. In 329this connection the large amount of work of this kind accomplished in the Sheffield Laboratory, at present under the guidance of Professor T. B. Johnson, should be mentioned.
It has happened that comparatively few publications on organic chemistry have appeared in the Journal, but it may be stated that the preparation of chloroform and its physiological effects were described by Guthrie (21, 64, 1832). Unknown to him, it had been prepared by Souberain, a French chemist, the previous year, but the former was the first to describe its physiological action. Silliman gave a sample to Doctor Eli Ives of the Yale Medical School, who used it to relieve a case of asthma. This was the first use of chloroform in medical practice (21, 405, 1832). Guthrie also described in the Journal (21, 284, 1832) his new process for converting potato starch into glucose, a method which is essentially the same as that used to-day in converting cornstarch into glucose. Lawrence Smith (43, 301, 1842 et seq.), Horsford (3, 369, 1847 et seq.), Sterry Hunt (7, 399, 1849), Carey Lea (26, 379, 1858 et seq.), Remsen (5, 179, 1873 et seq.), and others have contributed articles on organic chemistry.
Agricultural Chemistry.
Until near the middle of the nineteenth century, it was believed that plants, like animals, used organic matter for food, and depended chiefly upon the humus of the soil for their growth. This view was held even long after it was known that plant leaves absorb carbon dioxide and give off oxygen, and after the ashes of plants had been accurately analyzed.
This incorrect view was overthrown by the celebrated German chemist, Liebig, who made many investigations upon the subject, and, properly interpreting previous knowledge, published a book in 1840 upon the application of chemistry to agriculture and physiology in which he maintained that the nutritive materials of all green plants are inorganic substances, namely, carbon dioxide, water, ammonia (nitrates), sulphates, phosphates, silica, lime, magnesia, potash, iron, and sometimes common salt. He drew the vastly important conclusion that the effective 330fertilization of soils depends upon replenishing the inorganic substances that have been exhausted by the crops.
The fundamental principles set forth by Liebig have been confirmed, and it has been found that the fertilizing constituents most commonly lacking in soils are nitrogen compounds, phosphates, and potassium salts, so that these have formed the important constituents of artificial fertilizers. Liebig himself found that humus is valuable in soils, because it absorbs and retains the soluble salts.
The foundation established by Liebig in regard to artificial fertilizers has led to an enormous application of these materials, much to the advantage of the world’s food supply.
It was Liebig’s belief, in accordance with the prevailing views, that decay and putrefaction as well as alcoholic and other fermentations were spontaneous processes, and when the eminent French chemist, Pasteur, in 1857, explained fermentation as directly caused by yeast, an epoch-making discovery which led to the explanation of decay and putrefaction by bacterial action and to the germ-theory of disease, the explanation was violently opposed by Liebig and other German chemists. Pasteur’s view prevailed, however, and since that time it has been found that various kinds of bacteria are responsible for the formation of ammonia from nitrogenous organic matter and also for the change of ammonia into the nitrates that are available as plant-food.
The long-debated question as to the availability of atmospheric nitrogen for plant-food was settled in 1886 by the discovery of Hellriegel that bacteria contained in nodules on the roots, especially of leguminous plants, are capable of bringing nitrogen into combination and furnishing it to the plants.
No more than an allusion can be made to agricultural experiment stations where soils, fertilizers, foods and other products are examined, and where other problems connected with agriculture are studied.
The late S. W. Johnson of Yale studied with Liebig and subsequently did much service for agricultural chemistry in this country, by his investigations, his teaching, and his writings. His book, “How Crops Grow,” published 331in 1868, gave an excellent account of the principles of agricultural chemistry. He did much to bring about the establishment of agricultural experiment stations in this country, and for a long time he was the director of the Connecticut Station.
In the Journal, as early as 1827, Amos Eaton (12, 370) published a simple method for the mechanical analysis of soils to determine their suitability for wheat-culture, and Hilgard, between 1872 and 1874, described an elaborate study of soil-analysis. J. P. Norton, a Yale professor, in 1847 (3, 322) published an investigation on the analysis of the oat, which was awarded a prize of fifty sovereigns by a Scotch agricultural society, while Johnson, Atwater, and others have contributed articles on the analysis of various farm products.
Industrial Acids and Alkalies.
One hundred years ago sulphuric acid was manufactured on a comparatively very small scale in lead chambers. In 1818, an English manufacturer of the acid introduced the modern feature of using pyrites in the place of brimstone, while the Gay-Lussac tower in 1827 and the Glover tower in 1859 began to be applied as great improvements in the chamber process. Within about twenty years the contact process, employing platinized asbestos, has replaced the old chamber process to a large extent. It has the advantage of producing the concentrated acid, or the fuming acid, directly.
During our period the manufacture of sulphuric acid has increased enormously. Very large quantities of it have been used in connection with the Leblanc soda process in its rapid development. It came to be employed extensively for absorbing ammonia in the illuminating-gas industry, which was in its infancy one hundred years ago. New industries such as the manufacture of “superphosphates” as artificial fertilizers, the refining of petroleum, the manufacture of artificial dyestuffs and many other modern chemical products have greatly increased the demand for it, while its employment in the production of nitric and other acids, and for many other purposes not already mentioned, has been very great.
The manufacture of nitric acid has been greatly 332extended during our period on account of its employment for producing explosives, artificial dyestuffs, and for many other purposes. Chile saltpeter became available for making it about 1852. This acid has been manufactured recently from atmospheric nitrogen and oxygen by combining them by the aid of powerful electric discharges. This process has been used chiefly in Norway where water-power is abundant, as it requires a large expenditure of energy. A still more recent method for the production of nitric acid depends upon the oxidation of ammonia by air with the aid of a contact substance, such as platinized asbestos.
The production of ammonia, which was very small a hundred years ago, has been vastly increased in connection with the development of the illuminating-gas industry and the employment of by-product coke ovens. This substance is very extensively used in refrigerating machines and also in a great many chemical operations, including the Solvay soda process. Ammonium salts are of great importance also as fertilizers in agriculture. The conversion of atmospheric nitrogen into ammonia on a commercial scale is a recent achievement. It has been accomplished by heating calcium carbide, an electric-furnace product made from lime and coke, with nitrogen gas, thus producing calcium cyanamide, and then treating this cyanamide with water under proper conditions. Another method devised by Haber consists in directly combining nitrogen and hydrogen gases under high pressure with the aid of a contact substance.
Leblanc’s method for obtaining sodium carbonate from sodium chloride by first converting the latter into the sulphate by means of sulphuric acid and then heating the sulphate with lime and coal in a furnace was invented as early as 1791, but it was not rapidly developed and did not gain a foothold in England until 1826 on account of a high duty on salt up to that time. Afterwards the process flourished greatly in connection with the sulphuric acid industry upon which it depended, and with the bleaching-powder industry which utilized the hydrochloric acid incidentally produced by it, and, of course, in connection with soap manufacture and many other industries in which the soda itself was employed.
333About 1866 the Solvay process appeared as a rival to the Leblanc process. This depends upon the precipitation of sodium bicarbonate from salt solutions by means of carbon dioxide and ammonia, with the subsequent recovery of the ammonia. It has displaced the older process to a large extent, and it is carried on extensively in this country, for instance, at Syracuse, New York.
Other processes for soda depend upon the electrolysis of sodium chloride solutions. In this case caustic soda and chlorine are the direct products, and the chlorine thus produced and liquefied by pressure in steel cylinders, has become an important commercial article.
In earlier times wood-ashes were the source of potash and potassium salts. Wurtz in the Journal (10, 326, 1850) suggested the availability of New Jersey greensand as a source of potash and showed how this mineral could be decomposed, but it does not appear that this mineral has ever been utilized for the purpose. About 1861 the German potash-salt deposits began to be developed, and these have since become the chief source of this material. At present many efforts are being made to obtain potassium compounds from other sources, such as brines, cement-kiln dust, and feldspar and other minerals but thus far the results have not satisfied the demand.
Conclusion.
This account of chemical progress has given only a limited view of small portions of the subject, because the amount of available material is so vast in comparison with the space allowed for its presentation. Since the Journal has published comparatively little organic chemistry, it was decided to make room for a better presentation of other things by giving only a brief discussion of this exceedingly active and important branch of the science. For similar reasons industrial and metallurgical chemistry, and other branches besides, in spite of their great growth and importance, have been neglected, except for some incidental references to them, and some account of a few of the more important industrial chemicals.
It appears that we have much reason to be proud of the 334advances in chemistry that have been made during the Journal’s period, and of the part that the Journal has taken in connection with them, and there seems to be no doubt that this progress has not diminished during more recent times.
The present tendency of chemical research is evidently towards a still greater development of organic chemistry, and an increased application of physics and mathematics to chemical theory and practice.
The very great improvements that have been made in chemical education, both in the number of students and the quality of instruction, during the period under discussion, and particularly in rather recent times, gives promise for excellent future progress.
Note.
335