SPECIAL INVESTIGATIONS.
Reading edition
BLACK SAGE.
BOTANICAL DESCRIPTION AND DISTRIBUTION.
Ramona stachyoides (Benth.) Briquet (synonyms—Audibertia stachyoides Benth., Salvia mellifera Greene), commonly known as black sage (figs. 1 and 2), is a shrubby aromatic perennial, occurring from middle to southern California on low hills from April to June. The shrub attains a height of 3 to 6 feet and possesses herbaceous leafy branches with oblong leaves, green and wrinkled above and ash colored and hairy below. The flowers are white or lilac and in whorls or heads. The leaves have a strongly aromatic and decidedly camphoraceous odor, the woody branches being very brittle and also strongly aromatic.
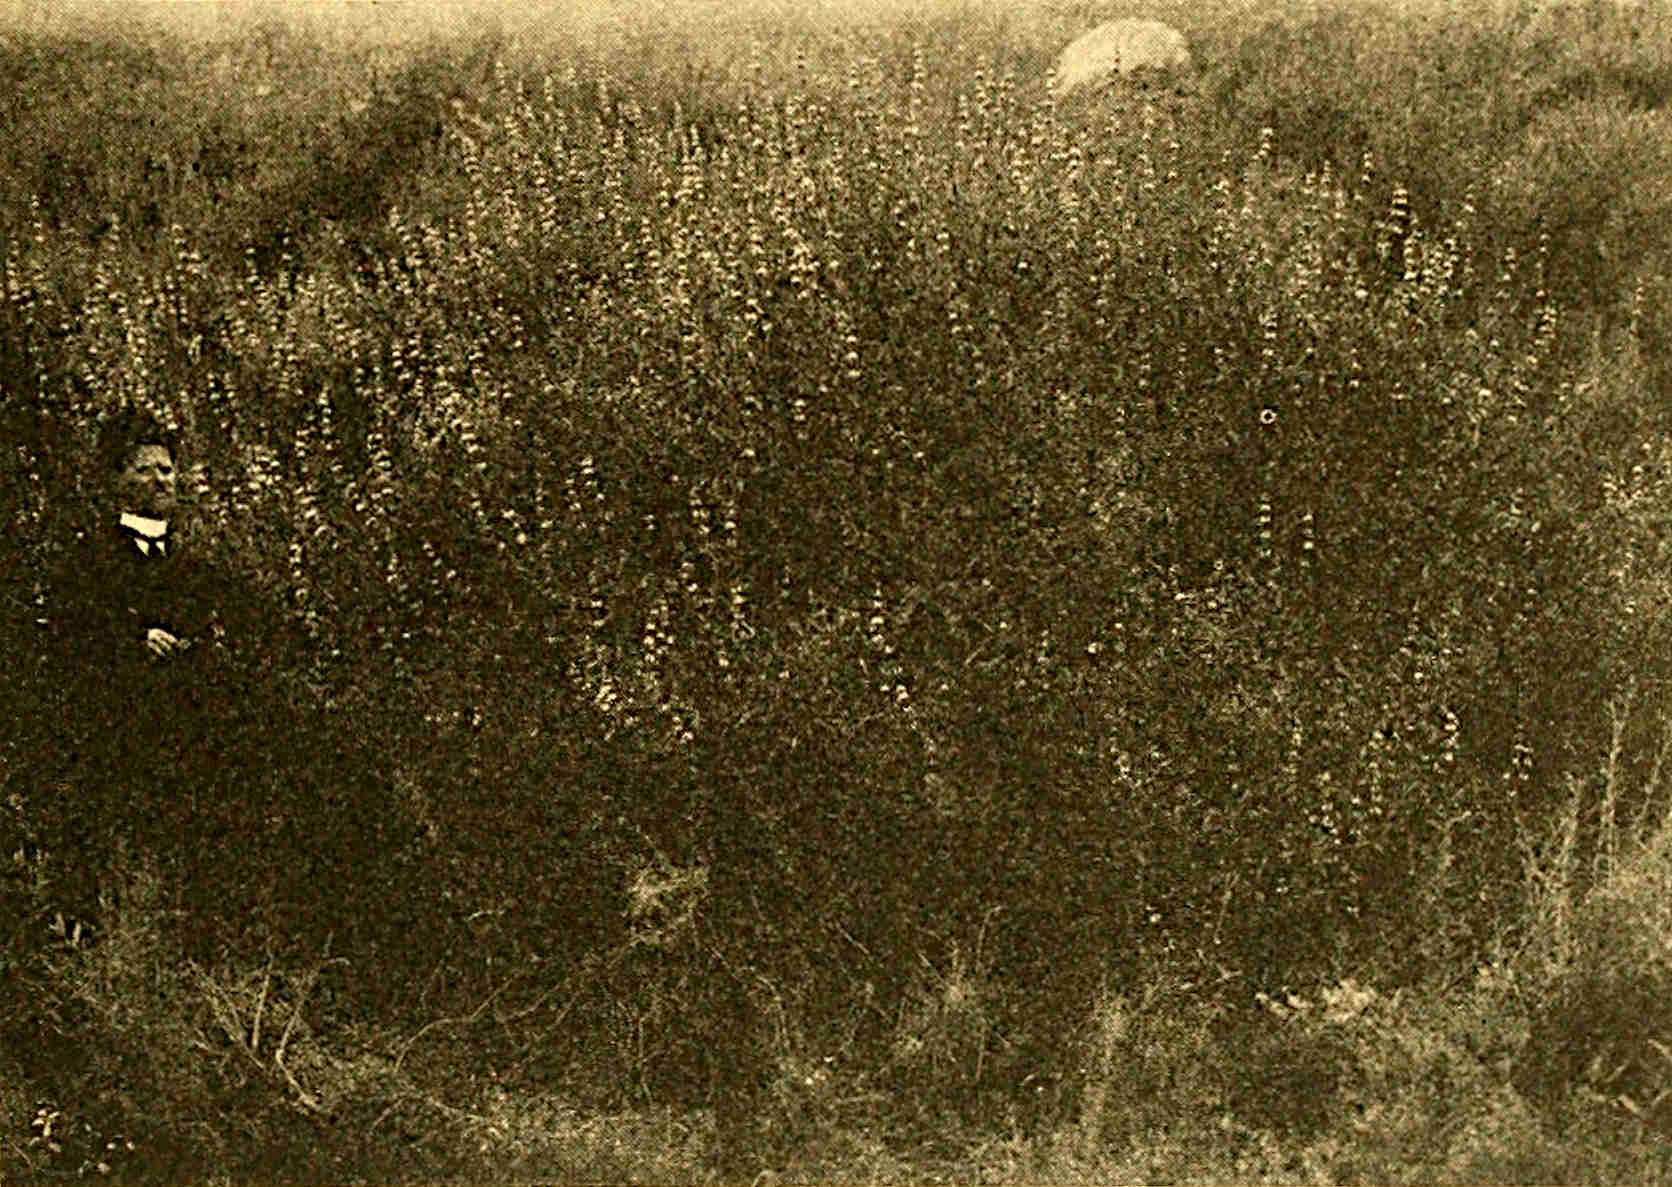
Fig. 1.—A plant of black sage (Ramona stachyoides) growing near Riverside, Cal.
DISTILLATION OF THE OIL.
A quantity of the fresh herb partly in bloom, including the flowering tops, branches, and leaves, was distilled by steam in the vicinity of Los Angeles, Cal., in April, 1908, and yielded 0.75 per cent of oil. The oil was nearly colorless and possessed a penetrating, camphoraceous, yet agreeable odor, with a bitter, camphorlike taste. At 24° C. the specific gravity was found to be 0.9144; specific rotation AD = +30.2°; re-fraction at 24° C., 1.4682. The oil was soluble with clear solution in 1½ volumes of 70 per cent alcohol, becoming turbid with 3½ volumes or over.
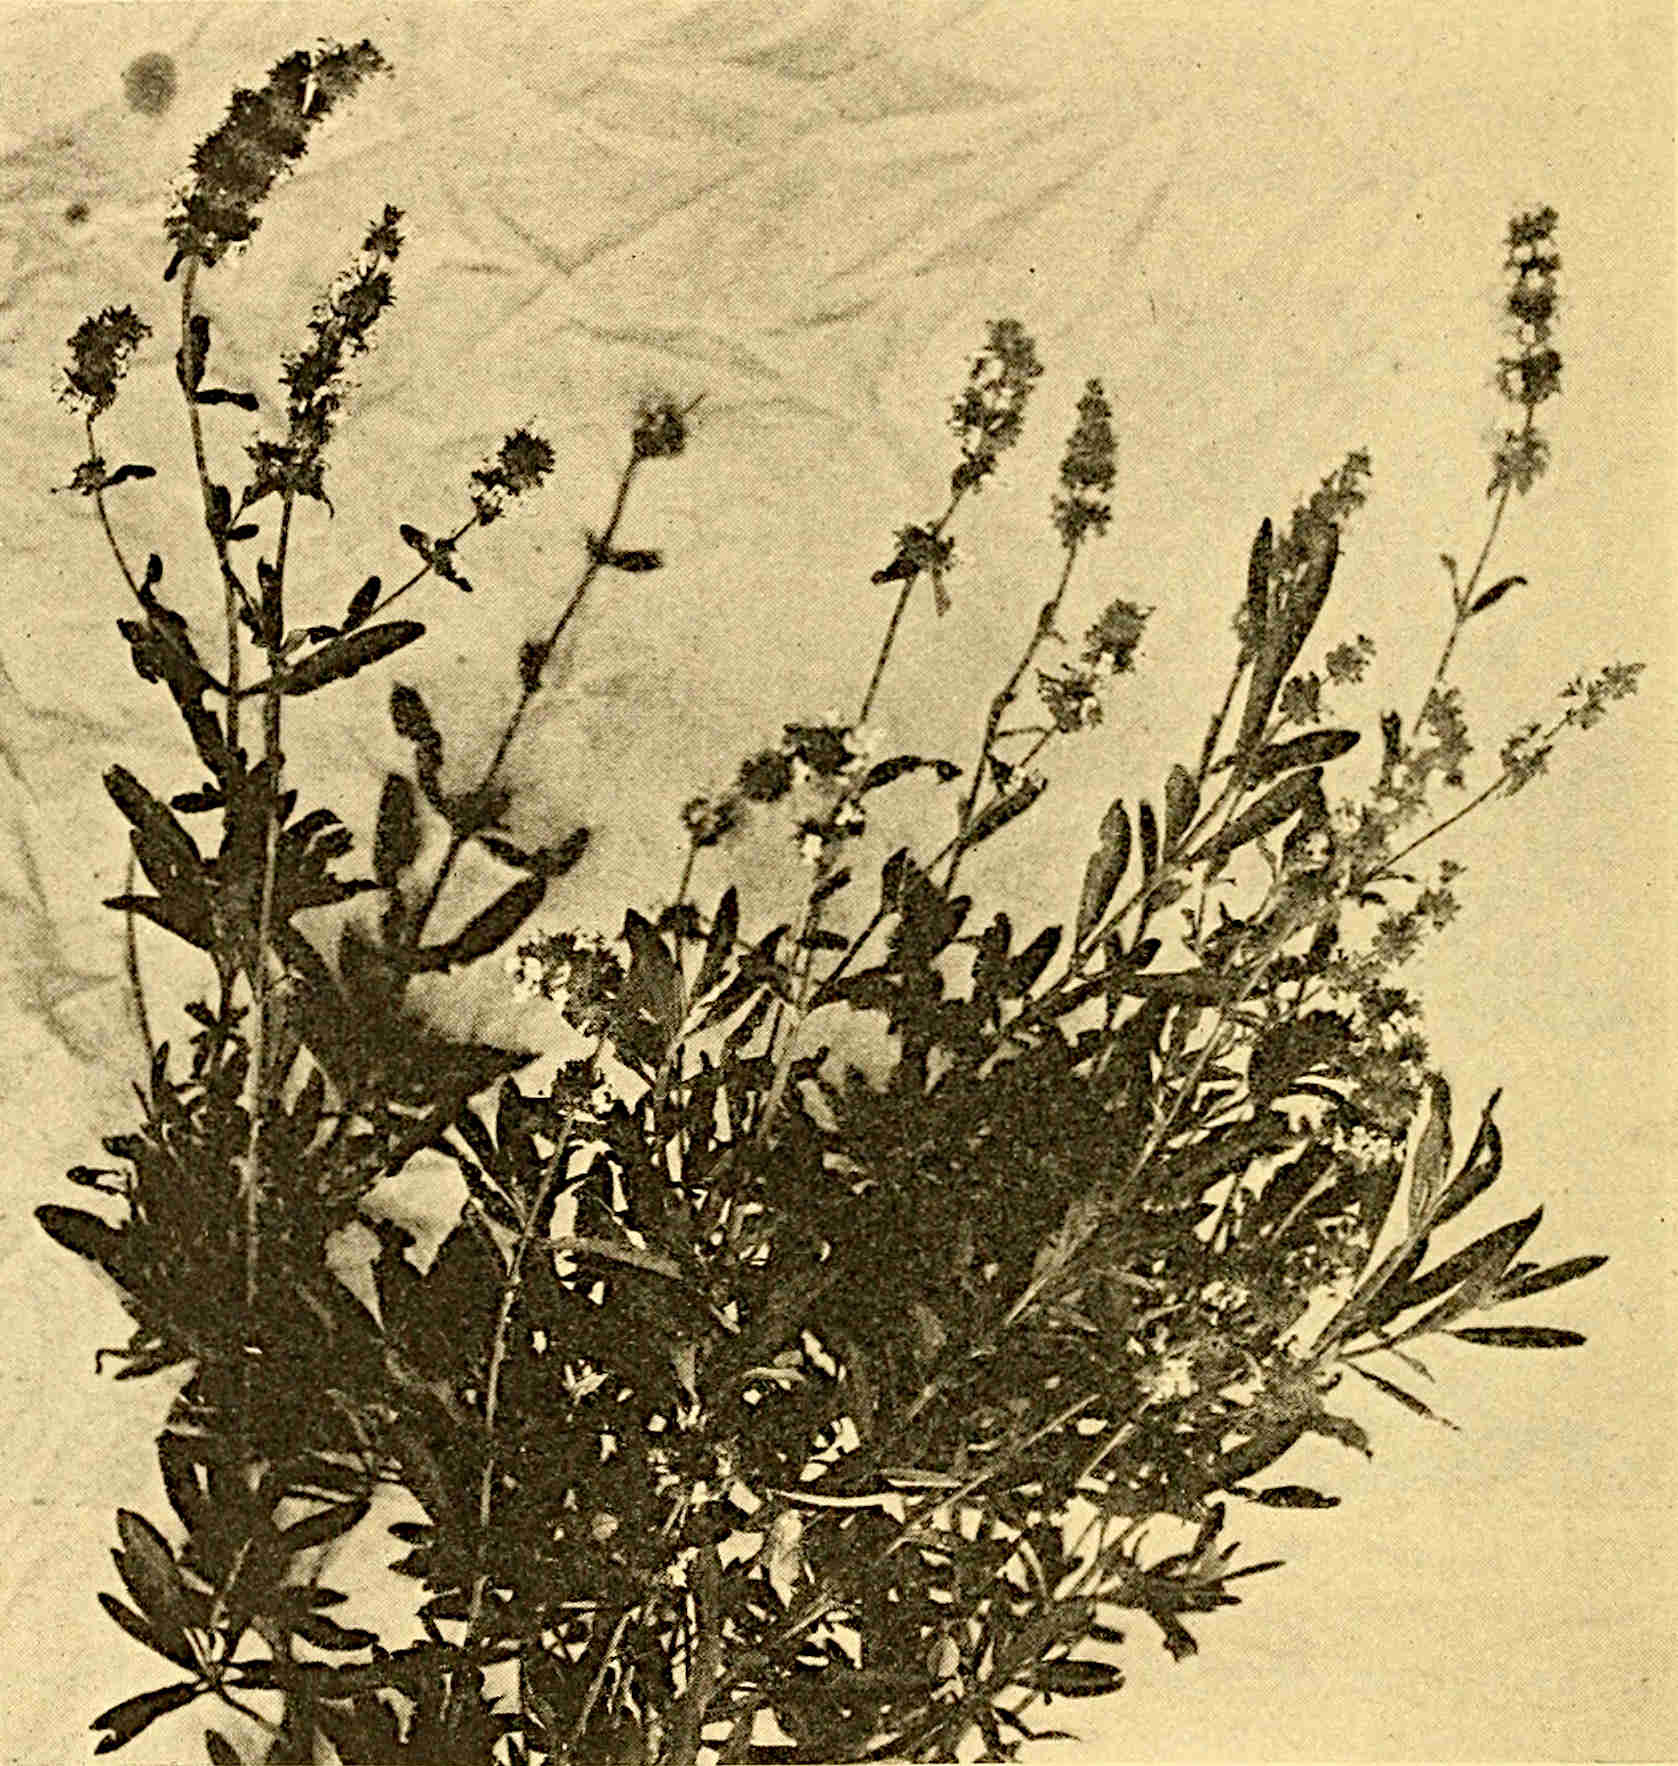
Fig. 2.—Flowering top of a plant of black sage.
SEPARATION OF STEAROPTENE.
Owing to the very strong camphoraceous odor of the oil, a separation of the stearoptene suggested itself. In order to separate a solid body which is held in solution by a volatile oil, the “freezing-out” method is usually employed. Accordingly 100 grams of the oil were subjected to a freezing mixture of ice and salt. A temperature of −15° C. was attained, and flaky crystals formed throughout the oil. The crystals were separated by being thrown on a force filter and the remaining oil again subjected to the cold, when a second lot was obtained, which was likewise separated. A total of 11.3 grams of crystals was separated, corresponding to a yield of 11.3 per cent. These crystals were soft and flaky in nature and possessed the characteristic odor of camphor.
IDENTIFICATION OF CAMPHOR.
In order to identify the crystalline substance obtained from the oil, a small quantity was sublimed, and the usual tests of melting point, boiling point, and rotation were applied. For further recognition of the compound, an attempt was made to prepare an oxime. Accordingly the method of Auwers[25] was applied, which, briefly, is as follows: To a solution of 10 parts of camphor in 10 to 20 times the amount of 90 per cent alcohol is added a solution of 7 to 10 parts of hydroxylamine hydrochlorid and 12 to 17 parts of a soda solution. If turbidity results, more alcohol is added and the mixture is heated on a water bath until a small portion of the solution remains clear upon the addition of water or until the resulting turbidity disappears, when a few drops of soda solution are added and no free camphor remains. The mixture is then diluted with water, filtered if necessary, and neutralized with dilute hydrochloric acid. The camphor oxime which separates is recrystallized from alcohol or ligroin. It melts at 118° to 119° C.
The above method applied to the sublimed crystals resulted in the formation of an oxime which melted at 120° to 124° C. Since an oxime was obtained (indicating possible ketonic characters), application was made of another reaction for ketones, namely, the formation of semicarbazone. Tiemann’s method[26] for the preparation of camphor semicarbazone was applied. The method is as follows: 1.5 grams of camphor dissolved in 2 cubic centimeters glacial acetic acid are treated with a solution of 1.2 grams of semicarbazid hydrochlorid and 1.5 grams of sodium acetate in 2 cubic centimeters of water. Water is added and the crystalline compound recrystallized from alcohol. The melting point of camphor semicarbazone is 236° to 238° C.
The sublimed crystals when treated in the above manner yielded a semicarbazone which melted at 232° C.
For a comparison of this substance with pure camphor, a tabulation was made of the more common physical properties and chemical tests.
Table I.—Comparison of properties of crystals from oil of black sage and of pure camphor.
Test. Crystals from oil of black sage. Crystals of pure camphor.
Melting point. 174° to 175° C. 175° C.
Boiling point. 205° C. 204° C.
Rotation in 50 mm. tube. +3.33° (20 per cent solution in alcohol). +3.51° (20 per cent solution in alcohol).
Oxime. M. p. 120° to 124° C. 118° to 119° C.
Semicarbazone. M. p. 232° to 233° C. 236° to 238° C.
The table shows very close similarities in the melting point, boiling point, and rotation of the crystals from the oil of black sage and of pure camphor. The melting points of the oximes and semicarbazones, though not corresponding so well, seemed to indicate that the crystals were in all probability camphor. To further confirm the assumption that the compound from the oil was camphor, an elementary analysis of the compound was made after being twice sublimed.
0.1273 gram of crystals gave 0.1199 gram H2O, corresponding to 10.5 per cent hydrogen.
0.1273 gram of crystals gave 0.3228 gram CO2, corresponding to 79.7 per cent carbon.
C10H16O camphor requires ⎰ ⎱ 79 per cent carbon. 10.5 per cent hydrogen.
0.1279 gram of crystals gave 0.1244 gram H2O, corresponding to 10.8 per cent hydrogen.
0.1279 gram of crystals gave 0.3761 gram CO2, corresponding to 79.9 per cent carbon dioxid.
The combustion results seemed to indicate that the compound is identical with that of camphor, as the above tabulation also clearly shows.
CHEMICAL EXAMINATION OF THE OIL.
CHEMICAL CONSTANTS.
Preliminary to the detailed chemical examination of the oil the usual chemical constants were determined.
By neutralization of a weighed quantity of the oil with standard potassium hydroxid V. S., the acid number (the number of milligrams of potassium hydrate required to neutralize 1 gram of oil) was found to be 2.
The ester number (the number of milligrams of potassium hydroxid required to saponify the esters in the oil) was found to be 2.5, which, calculated as bornyl acetate, corresponds to 0.88 per cent.
The ester number after acetylization of the saponified oil with acetic anhydrid (and which represents the total amount of alcohols present) was 27.1, which, calculated as borneol, represents a total of 7.58 per cent of borneol in the oil, both free and in combination.
FREE ACIDS.
The original oil was slightly acid, as indicated by the acid number previously mentioned. The free acid was shaken out from a quantity of the oil with a 10 per cent solution of sodium carbonate. The shaking was repeated several times and the alkaline liquids united. The united alkaline liquids were shaken out with ether in order to remove any oil held in suspension. The sodium-carbonate solution was then evaporated to a small bulk on a water bath, acidified with sulphuric acid, and distilled with steam. No oily globules separated, showing absence of higher insoluble acids. The distillate, which was decidedly acid, was neutralized with sodium-carbonate solution and evaporated to a small volume. The liquid which now represented the sodium salts of the free acids present in the oil was precipitated fractionally with a dilute silver-nitrate solution. Four fractions resulted. Each fraction was dried to constant weight and burned.
Fraction 1. 0.1014 gram silver salt = 0.0785 gram silver = 76.3 per cent silver.
Fraction 2. 0.1000 gram silver salt = 0.077 gram silver = 77 per cent silver.
Fraction 3. 0.1116 gram silver salt = 0.0859 gram silver = 76.9 per cent silver.
Fraction 4. 0.1088 gram silver salt = 0.077 gram silver = 70.8 per cent silver.
Fraction 4 indicates the presence of formic acid, the silver salt of which requires, theoretically, 70.5 per cent of silver. Fractions 1, 2, and 3 indicate silver carbonate (which requires, theoretically, 78 per cent of silver) with a slight admixture of silver formate. The presence of silver carbonate was caused by a possible slight excess of sodium carbonate being added when the acid distillate was neutralized.
COMBINED ACIDS.
Saponification.—For the purpose of determining the acids held in combination in the oil in the form of esters, the oil was saponified with alcoholic potassium hydrate by heating on a water bath with a reflux condenser for one-half hour. Water was added to the mixture, and the oil separated in a layer. After removing the excess alcohol on a water bath, the alkaline solution was shaken out with ether to remove any adhering oil. The remaining solution was evaporated to a small volume, acidified with sulphuric acid, and distilled with steam.
The distillate from the above was extracted with ether and the ether evaporated spontaneously. Only a trace of an acid residue remained, which was neutralized with a solution of potassium hydroxid and precipitated in three fractions:
Fraction 1. 0.1012 gram silver salt = 0.0893 gram silver = 88 per cent silver.
Fraction 2. 0.0774 gram silver salt = 0.0637 gram silver = 82.3 per cent silver.
Fraction 3. 0.0758 gram silver salt = 0.0502 gram silver = 66.2 per cent silver.
The first two precipitates, when dried, consisted principally of silver oxid, which, theoretically, contains 89.2 per cent of silver. A slight excess of potassium hydroxid during neutralization was doubtless responsible. Fraction 3 would seem to point to the presence of acetic acid in the oil, silver acetate requiring 64.6 per cent of silver.
The aqueous acid portion remaining after the ether extraction was neutralized with sodium carbonate concentrated to small bulk and precipitated with silver nitrate in three fractions. Fraction 1 contained 76.2 per cent of silver; fraction 2, 77 per cent; and fraction 3, 74 per cent. Since silver formate contains 70.5 per cent of silver, a trace of formic acid is possibly present in the oil in combination.
The esters of the oil, as shown by the above results, are present in the oil principally as acetates, with a possible trace of formates.
FRACTIONATION OF THE OIL.
In order to ascertain the total percentage of camphor and to separate the remaining constituents as completely as possible, a quantity of the oil was fractionated into seven fractions, as follows:
Fraction 1, 160° C.; fraction 2, 160° to 170° C.; fraction 3, 170° to 178° C.; fraction 4, 178° to 182° C.; fraction 5, 182° to 186° C.; fraction 6, 186° to 190° C.; fraction 7, 190° to 195° C. These fractions (125 grams) were refractionated into 10 separate fractions, as shown in Table II, a determination of the physical properties of each fraction also being made.
Table II.—Fractionation of the oil of black sage, showing the physical properties of the fractions.
Fraction. Temperature. Distilled over. Specific gravity at 26° C. Rotation Re-fraction ND 28° C. Remarks.
Degrees C. Per cent. Degrees.
1 Below 160 2.5 0.8070 +6.9 1.4570 Slight terebinthine odor.
2 160 to 170 6.8 .8768 +10.1 1.4613 Cineol-like odor.
3 170 to 174 7.8 .8865 +10.1 1.4640 Do.
4 174 to 178 12.1 .8920 +10 1.4648 Decidedly cineol-like odor.
5 178 to 182 14.8 .8996 +10.2 1.4652 Do.
6 182 to 186 8.6 .9077 +11.5 1.4659 Slight camphoraceous odor.
7 186 to 190 8 .9105 +11.1 1.4673 Strong camphoraceous odor.
8 190 to 195 8.1 .9130 +11.7 1.4683 Do.
9 195 to 200 7.7 .9170 +11.6 1.4710 Do.
10 200 to 208 11.6 .9220 +10.4 1.4710 Do.
Residue 208 and above 12 .9236 1.4854 Do.
IDENTIFICATION AND SEPARATION OF THE CONSTITUENTS.
Pinene.—The first fraction distilling below 160° C., and which possessed an odor of turpentine, was tested for pinene by means of the nitrochlorid reaction.[27] A deep blue coloration was obtained with slight turbidity, indicating a possible trace of pinene.
Cineol, or eucalyptol.—Tests were made in fractions 2, 3, 4, 5, and 6 for cineol, which was easily recognized by its odor. For a qualitative test the iodol reaction was used, crystals of cineol iodol which melted at 111° to 112° C. forming in each fraction. Fractions 3, 4, and 5, which smelled strongly of cineol and which doubtless contained the major portion of cineol in the oil, were assayed by means of the phosphoric acid method, as directed in the United States Pharmacopœia for 1900.[28] From these four fractions a total amount of 22.5 per cent of cineol was obtained, calculated from the original oil. This figure represents approximately the percentage of cineol in the oil, although it is low rather than high, since fractions 2 and 6 both showed the presence of cineol by qualitative tests, but the quantitative estimation in these fractions was impossible owing to the preponderance of other constituents in the fractions.
A test for terpinene in fraction 6, by means of the terpinene nitrosite reaction, produced a characteristic blue coloration, but the crystalline nitrosite would not separate.
Camphor.—A strong odor of camphor being distinguishable in fractions 7, 8, 9, 10, and in residue, a quantitative separation was made as completely as possible by means of the “freezing-out” method. Between 186° and 190° C. some crystals of camphor began to form in the inner tube of the condenser, and at 195° C. the condenser had to be kept jacketed with steam to prevent clogging, so rapidly did the camphor distill over. The fractions above 195° C. were practically solid. The camphor which separated at ordinary temperature was filtered on a force filter, and the liquid portion of the fractions subjected to freezing successively until camphor no longer separated. It is apparent that the separation of the camphor from these small fractions by freezing out is rather inaccurate because of the losses in transferring and filtering. From the above fractions, however, a quantity of camphor was obtained corresponding to about 40 per cent of the original oil. This figure is low, for the separation on a larger scale working with much larger fractions would reduce to a considerable degree the loss of camphor which is unavoidable in such small fractions.
The fractions distilling between 195° and 208° C. yielded crystals when treated with bromin in a petroleum-ether solution of the oil. The crystals melted at 130° C. Thujone tribromid melts at 122° C. A trace of thujone is therefore probably present in the oil. It is very possible, in view of the fact that the acetylization of the oil disclosed some free alcohol, that the last fraction contained some borneol, which boils at 212° C.
SUMMARY.
The results of the experiments would seem to indicate that the oil of black sage is composed essentially of camphor (more than 40 per cent) and cineol (22.5 per cent), with a small quantity of an alcohol, probably borneol, both free and as an ester, and a small quantity of the ketone thujone, with traces of the terpenes pinene and terpinene. Free formic acid was found, and only traces of combined acetic and formic acids in the form of esters.
The constituents of possible economic importance in the oil are camphor and cineol, both of which possess considerable medicinal value, the former being used also very extensively in the arts. These constituents, possessing strong antiseptic virtues, no doubt impart antiseptic properties to the oil. Inasmuch as the yield of oil from the fresh herb approximates 1 per cent, if distilled during the full flowering stage, and furthermore, since the plant thrives on low sandy hills or wastes, it is very probable that the shrub could be grown profitably both for its oil and for the large amount of camphor and cineol capable of being isolated from it.
WILD SAGE.
BOTANICAL DESCRIPTION AND DISTRIBUTION.
Artemisia frigida Willd., commonly known as wild sage, mountain sage, pasture sagebrush, and wormwood sage (figs. 3 and 4), is a hardy perennial 6 to 20 inches high, with a woody base and white silky leaves. The numerous yellow flowers, arranged in a racemelike head, possess a strongly camphoraceous odor. The leaves are also strongly aromatic. The plant abounds on dry sandy hilltops from the Dakotas west to Idaho, north into Canada, and as far south as Texas.
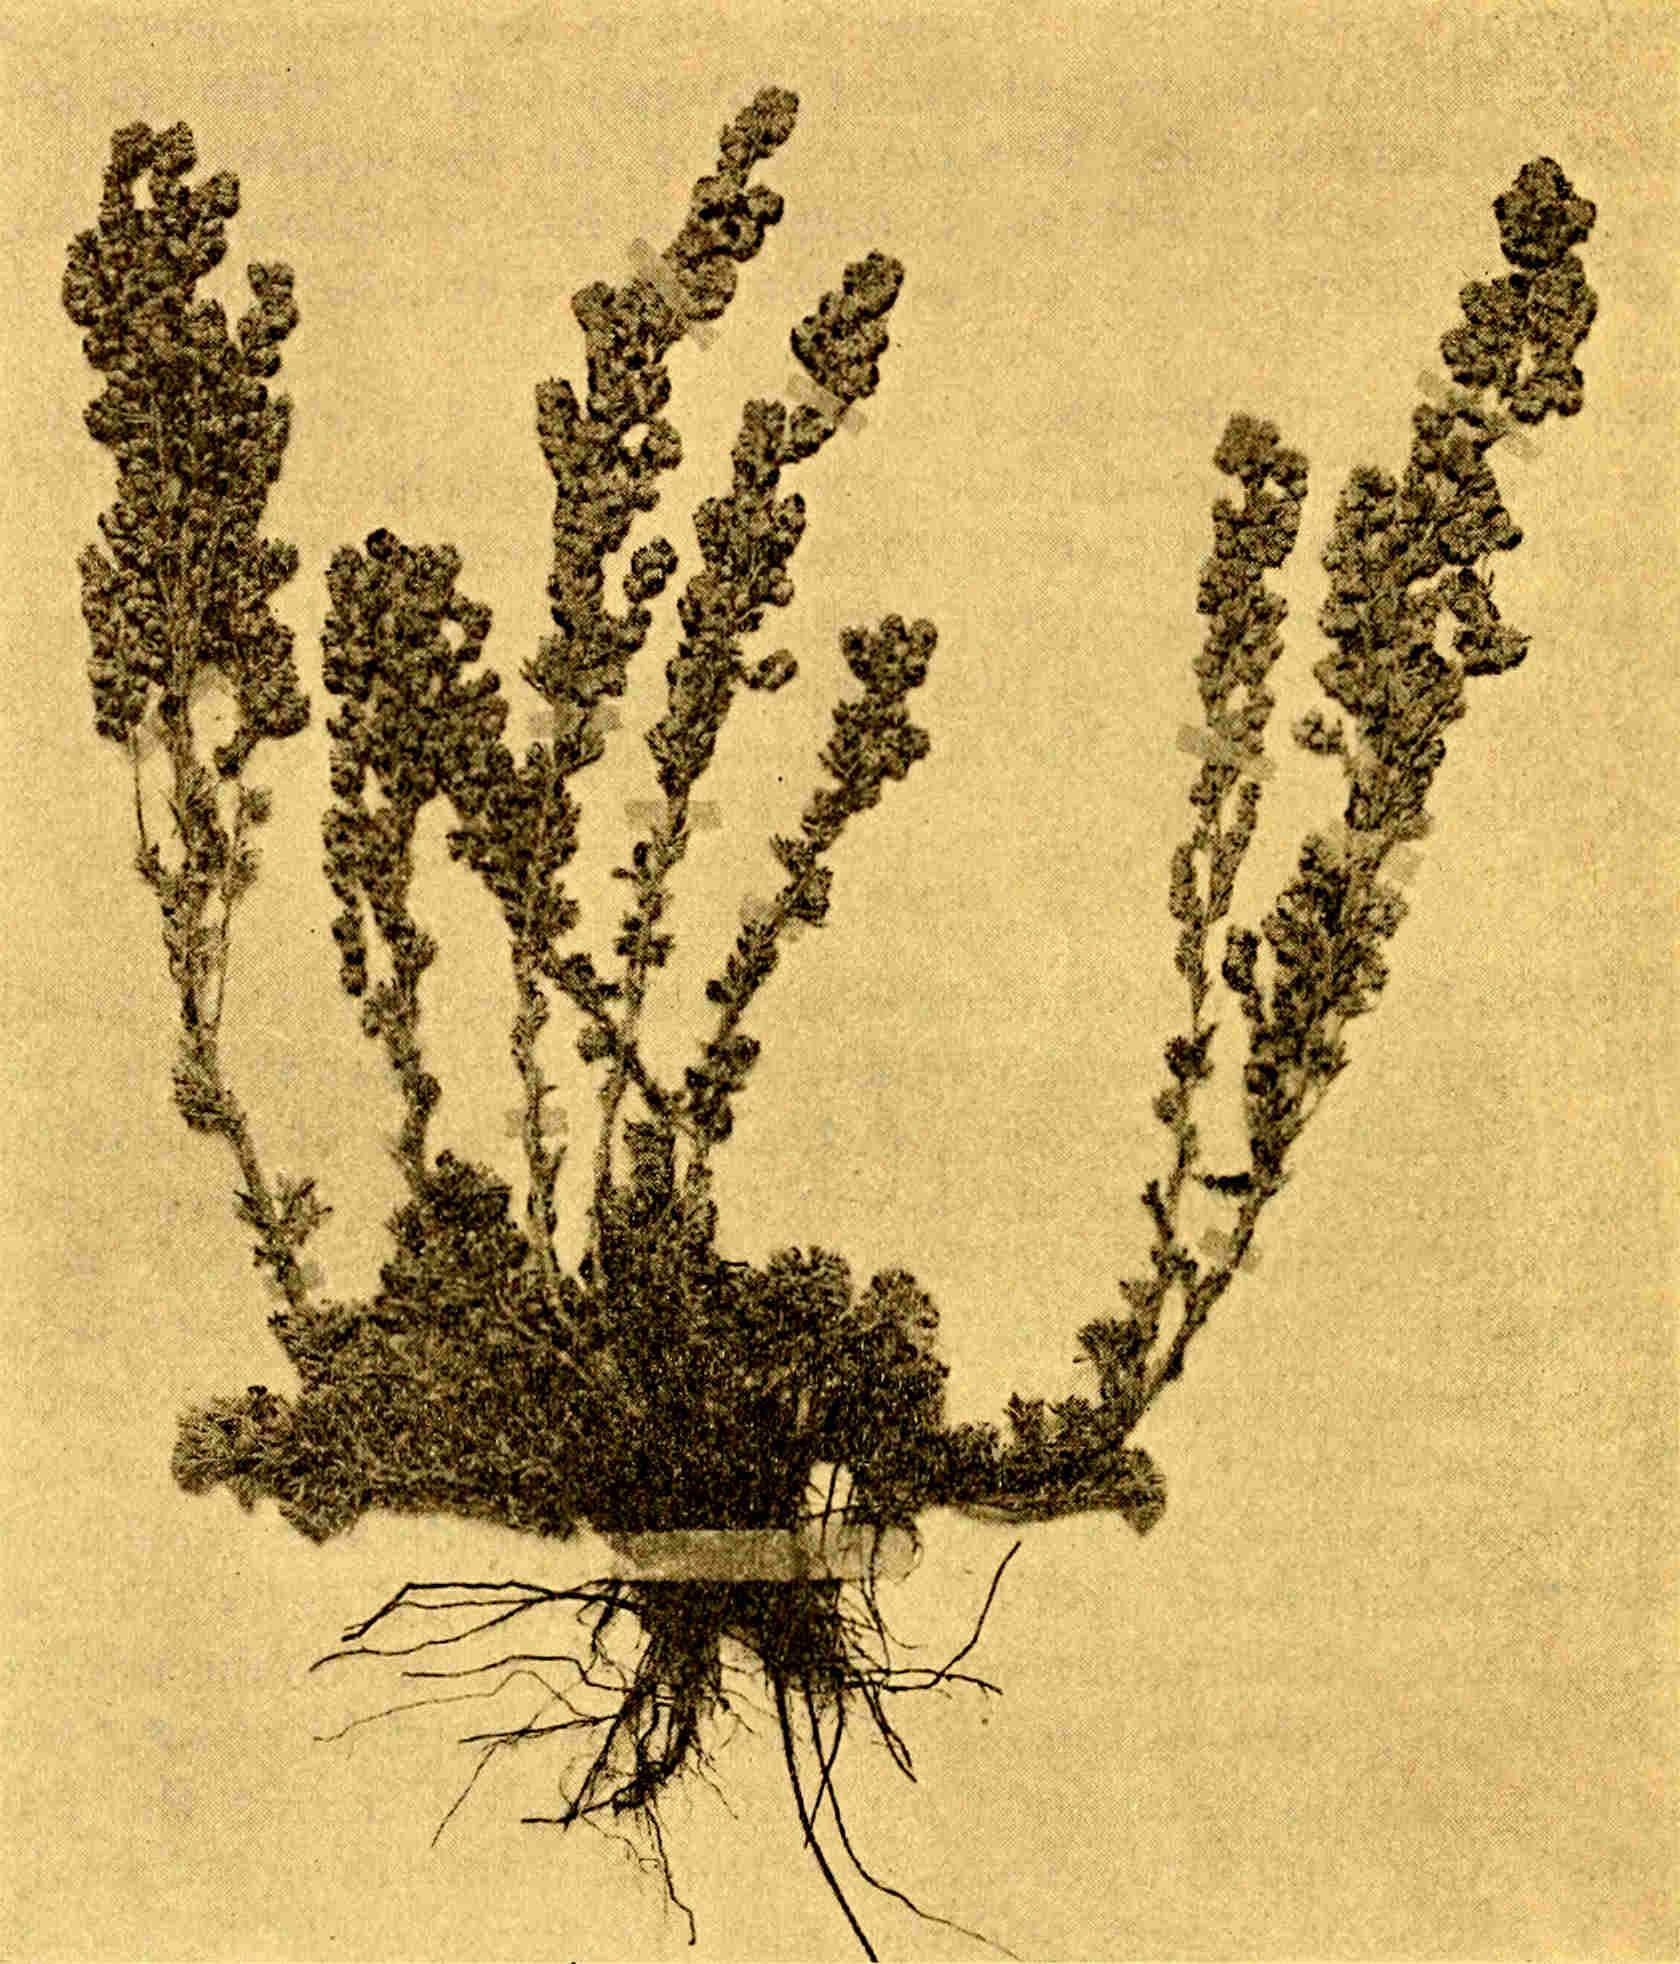
Fig. 3.—A plant of wild sage (Artemisia frigida).
DISTILLATION OF THE OIL.
The oil distilled from wild sage was briefly reported by the writer in 1905[29] and 1906.[30] The promising preliminary results encouraged a further investigation of this plant. During the summers of 1907 and 1908 larger quantities of this interesting wild plant were distilled in South Dakota, a yield of 0.26 per cent of a very fragrant essential oil being obtained from plants which had passed their flowering stage. When the plant is distilled during its flowering stage the yield of oil is about 0.41 per cent.
The oil obtained by the distillation of the whole plant was beautiful pale green in color, with an agreeable fatty and camphoraceous odor and a slightly bitter camphorlike taste. The specific gravity of the oil at 24° was 0.940; specific rotation AD = −24.2°; re-fraction ND 24°, 1.4716. The oil was soluble in 1 volume of 80 per cent alcohol, becoming turbid in 2 volumes or over.
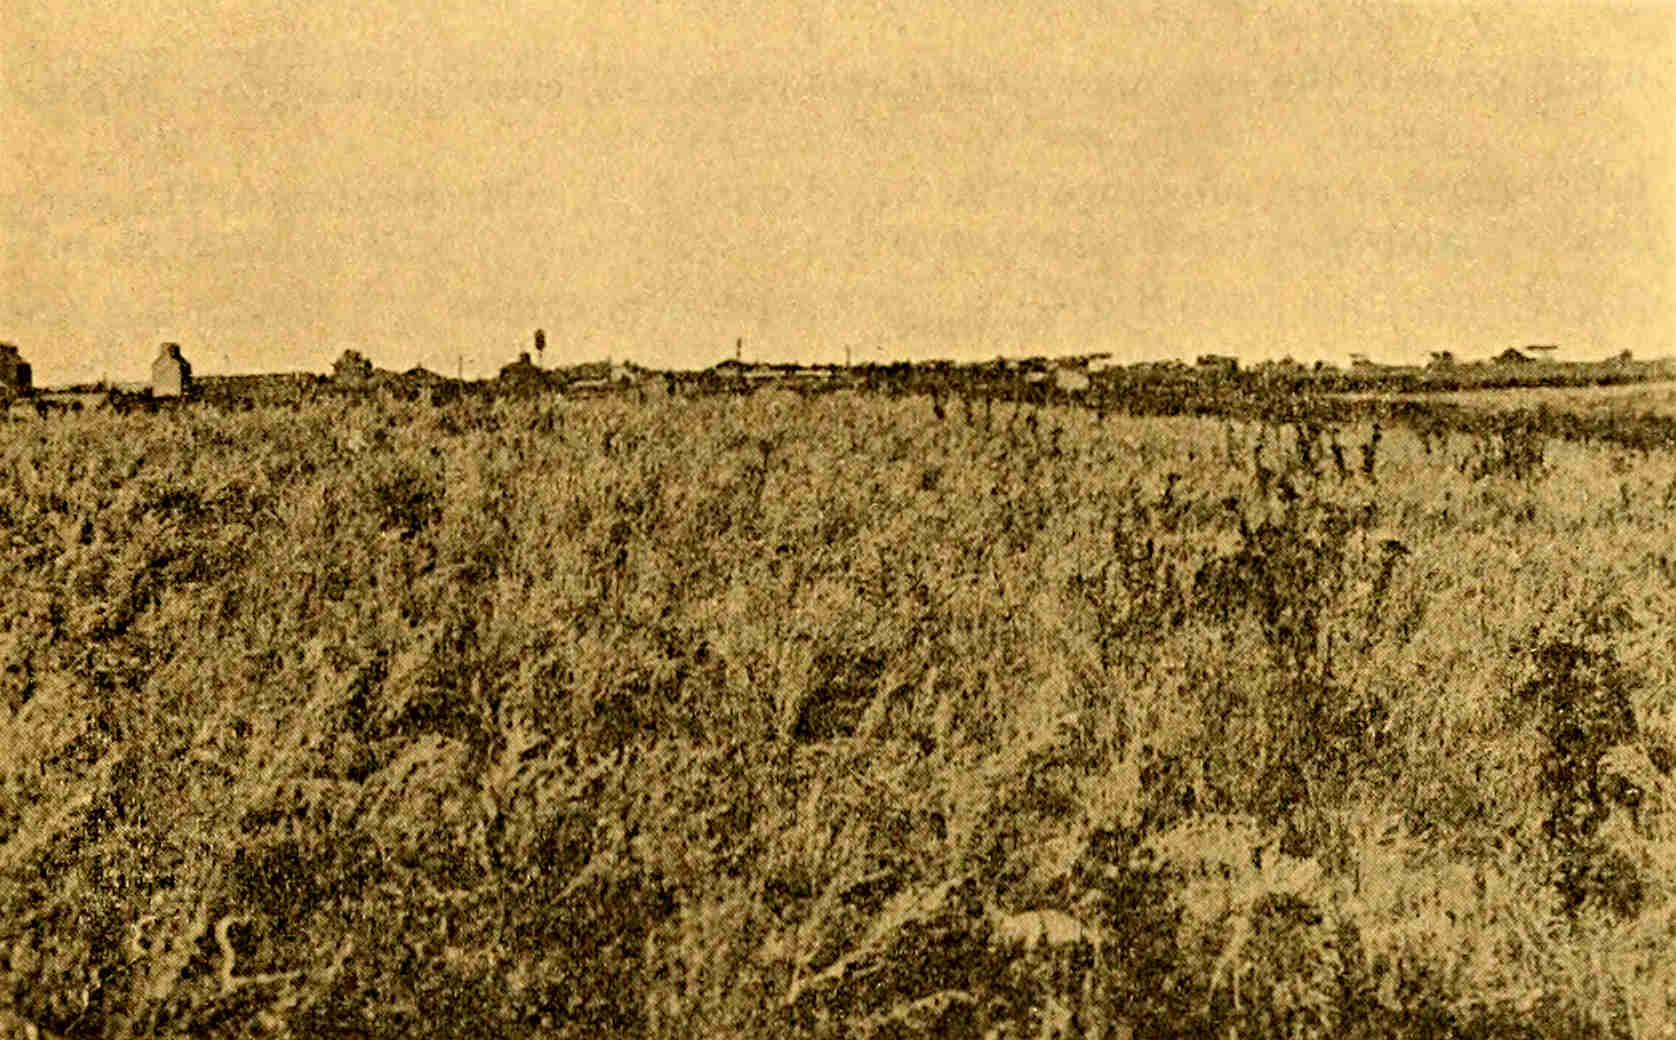
Fig. 4.—A field of wild sage near Webster, S. Dak.
SEPARATION OF STEAROPTENE.
During the distillation and filtration of the oil, small crystals were observed at the mouth of the distillation apparatus and also at the mouth of the funnel after standing over night. In order to separate this stearoptene (solid portion of the oil) from the elaoptene (liquid portion) 50 grams of the oil were subjected to a freezing mixture of ice and salt for several hours. As a result crystals separated in the form of white flakes. The crystals were thrown into a force filter and weighed, a total of 3 per cent resulting.
IDENTIFICATION OF CRYSTALLINE COMPOUND.
After recrystallization of the above crystals from alcohol the properties of the crystals compared very favorably with levo borneol, as shown in Table III.
Table III.—Comparison of properties of crystals from oil of wild sage and of pure borneol.
Test. Crystals from oil of wild sage. Crystals of pure borneol.
Color. White. White.
Odor. Camphorlike. Camphorlike.
Taste. Bitter, camphorlike. Bitter, camphorlike.
Boiling point. 210° to 215° C. 212° C.
Melting point. 203° C. 203° to 204° C.
Specific rotation. −32°. −37°.
To further confirm the above results, which seemed to indicate that the compound was identical with levo borneol, an elementary analysis was made.
0.1237 gram of the substance gave 0.3499 gram CO2, corresponding to 77.2 per cent carbon.
0.1237 gram of the substance gave 0.1252 gram H2O, corresponding to 11.4 per cent hydrogen.
C10H18O borneol requires ⎰ ⎱ 77.8 per cent carbon. 11.7 per cent hydrogen.
The elementary composition substantiates the assumption that the crystals are identical with levo borneol.
CHEMICAL EXAMINATION OF THE OIL.
CHEMICAL CONSTANTS.
The usual chemical constants were determined, namely, the acid number, ester number, saponification number, and acetylization number.
The acid number, denoting the amount of free acids contained in the oil and expressed in milligrams of potassium hydroxid, was determined by simple neutralization of the oil with standard potassium hydrate volumetric solution.
The ester number, denoting the amount of esters (combination of alcohols and acids) in the oil and expressed in milligrams of potassium hydroxid, was determined by saponification of the ester compounds with alcoholic potassium hydrate.
The acetylization number, or the ester number determined after acetylization of the oil with acetic anhydrid, signifies the total amount of alcohol or alcohols in the oil.
The constants of the oil were determined with the following results:
Acid number, 2.5, calculated as acetic acid, indicates 0.26 per cent acetic acid.
Ester number, 25, calculated as bornyl acetate, indicates 8.7 per cent bornyl acetate, which is equivalent to 6.8 per cent of free borneol.
Saponification number, 27.5.
Acetylization number, 139, corresponds to 42.67 per cent of total borneol in the oil, or, deducting the 6.8 per cent of free borneol as the ester, to 38 per cent of free borneol.
Assuming that the stearoptene obtained was borneol, a determination of the constants of the stearopteneless oil was made. The acid number remained practically the same, being 2.3; the ester number differed only very slightly, being 24.7; but the acetylization value obtained was only 132, which corresponded to but 40 per cent of total borneol. This is in strict conformity with the assumption, which seemed to be sufficiently proved, that the stearoptene separated from the oil by freezing was borneol. The stearopteneless oil was nearly 3 per cent poorer in borneol than the original oil, as shown above. It is to be remembered that 3 per cent of crystalline borneol was removed by freezing the original oil, hence the lowering of the borneol content of the stearopteneless oil.
FREE ACIDS.
The determination of the free acids was accomplished by repeatedly shaking a portion of the original oil with a 10 per cent sodium carbonate solution. After removing the adhering oil from the alkaline liquid by shaking with ether, the solution was acidified and distilled with a current of steam. A few oily globules floated on the surface of the liquid. These were extracted with ether and the ether evaporated. A small amount of oily residue remained, which was distinctly acid. The oily residue was exactly neutralized with sodium hydrate and precipitated with silver nitrate solution in three fractions:
Fraction 1. 0.0377 gram silver salt gave 0.0162 gram silver = 42.9 per cent silver.
Fraction 2. 0.0206 gram silver salt gave 0.0091 gram silver = 44.1 per cent silver.
Fraction 3. 0.0663 gram silver salt gave 0.3000 gram silver = 45.2 per cent silver.
The three fractions appear to be a mixture of caprylic and œnanthylic acids. Silver caprylate requires 42.9 per cent silver; silver œnanthylate requires 45.5 per cent silver.
A small amount of the insoluble free acids was therefore caprylic acid (octoic acid), the major portion being œnanthylic acid (heptoic acid).
The distillate from which the oily acids were extracted by ether was still slightly acid and was accordingly neutralized with sodium carbonate and precipitated with silver nitrate, two fractions being obtained. The first corresponded to silver carbonate, due to a slight excess of sodium carbonate; the second, only trifling in quantity, indicated the presence of only a trace of formic acid in the free condition.
Œnanthylic, or heptoic, acid seems to be the predominating free acid in the oil, with slight traces of formic and caprylic, or octoic, acids.
COMBINED ACIDS.
The esters in the oil, being combinations of alcohols and acids, serve as a basis for the identification of the acids in combination. In order to accomplish a separation of the combined acids a small quantity of the oil was saponified with alcoholic potassium hydroxid by heating on a water bath for half an hour. After dilution of the mixture with water and separation of the oil the alkaline liquid, which contained a small amount of the oil held in suspension, was shaken out with ether. The liquid was then acidified with sulphuric acid and distilled with steam. The oily globules which separated on the distillate were extracted with ether and the solvent evaporated. A small amount of an oily liquid with very offensive odor remained. This mixture of oily acids was neutralized with sodium hydrate and precipitated with a dilute solution of silver nitrate. Two fractions resulted:
Fraction 1. 0.0430 gram silver salt gave 0.0160 gram silver = 37.2 per cent silver.
Fraction 2. 0.0440 gram silver salt gave 0.0197 gram silver = 44.7 per cent silver.
From the results obtained it is evident that the fractions consist of the silver salt of undecylic acid, which requires theoretically 36.8 per cent of silver, and silver salt of heptoic (œnanthylic) acid, which requires 45.5 per cent of silver.
The aqueous distillate from the above, after being made neutral with sodium carbonate, was evaporated to small volume and precipitated in three fractions with silver nitrate:
Fraction 1. 0.243 gram silver salt gave 0.1837 gram silver = 75 per cent silver.
Fraction 2. 0.3255 gram silver salt gave 0.2370 gram silver = 72.9 per cent silver.
Fraction 3. 0.3492 gram silver salt gave 0.1840 gram silver = 52.9 per cent silver.
The greater portion of the soluble combined acids consisted of valerianic acid, the silver salt requiring 51.6 per cent of silver. A trace of formic acid was also indicated in combination as an ester in fraction 2, above.
The chief acids in combination as esters in the oil appear to be œnanthylic (heptoic) and valerianic, the former being preponderant. Formic and undecylic acids occur only as traces. All of the above are no doubt combined in the oil as esters of borneol.
FRACTIONATION OF THE VOLATILE OIL.
One hundred grams of the original oil were subjected to fractionation and separated into six fractions of 5 degrees each, beginning with 175° C. Those fractions together with the residue were again fractionated in order to insure a better separation of the constituents.
Table IV.—Fractionation of oil of wild sage, showing the physical and chemical properties of the fractions.
Fraction. Temperature. Distilled. Specific gravity at 24° C. Rotation in 50 mm. tube. Ester number. Remarks.
Degrees C. Per cent. Degrees.
1 Below 175[31] 13.6 0.9084 −9.5 2.6 Eucalyptuslike (cineol) odor.
2 175 to 180 14.0 .9196 −8.1 6.7 Do.
3 180 to 185 6.0 .9269 −7.6 8.9 Do.
4 185 to 190 4.5 .9313 −6.4 12.0 Slightly camphoraceous odor.
5 190 to 195 10.0 .9401 −8.6 25.8 Camphoraceous odor.
6 195 to 205 9.5 .9478 −9.7 34.8 Decidedly camphoraceous; free borneol crystallized in condenser.
7 205 to 215 9.0 .9562 −10.6 56.4 Fraction almost solid (borneol).
8 215 to 230 9.5 .9600 −10.8 75.2 Fraction partially solidified.
9 230 to 245 3.5 .9570 −5.8 70.7 Few crystals separated.
10 245 and above 9.0 .9830 47.0 Dark, sirupy, camphoraceous.
IDENTIFICATION AND SEPARATION OF THE CONSTITUENTS.
Cineol.—Fraction 1, 175° C., possessed a strong eucalyptuslike odor and was tested for cineol by means of iodol. The tetraiodopyrol (iodol) addition product of cineol formed into well-defined, nearly colorless crystals, melting at 110° to 113° C. This crystalline addition product of iodol formed in the first four fractions; in fraction 5, however, only a trace of crystals appeared.
The presence of cineol having been proved, a quantitative estimation of the compound was made in fractions 1, 2, 3, and 4. Fraction 5, which contained only a very small quantity of cineol, did not admit of estimation by the phosphoric-acid method, which is reliable only when large percentages of cineol are present.
The fractions yielded the following percentages of cineol: 1, 40 per cent; 2, 70 per cent; 3 and 4, 43.7 per cent. Calculating from the original oil as a basis, the above results correspond to 19.7 per cent of cineol in the original oil.
Fenchone.—Fraction 5, boiling from 190° to 195° C., was a heavy liquid with a strong camphorlike odor. Pure levo fenchone[32] from thuja oil is an oily liquid with a strong camphoraceous odor; boiling at 192° to 194° C.; specific gravity at 19° C., 0.946; (AD) −66.9°.
An oxime was prepared from the fraction by reaction with hydroxylamine hydrochlorid according to the method of Wallach,[33] which is as follows: To 5 grams of fenchone dissolved in 80 cubic centimeters of absolute alcohol is added a solution of 11 grams of hydroxylamine hydrochlorid in 11 grams of hot water. Six grams of powdered potash are added. The oxime separates in the form of crystals, upon standing for some time. Recrystallized from alcohol it melts at 164° to 165° C.
The oxime formed from the fraction by the above method, after recrystallization from ethyl acetate, melted at 170° C.
Provided that fraction 190° to 195° C. consists chiefly of fenchone the oil should contain 8 to 10 per cent of this compound.
Borneol.—The total amount of borneol contained in the oil was determined by the saponification of a small quantity of the original oil and subsequently fractionating the saponified oil. Twenty-five grams of the saponified oil were carefully fractionated and then refrigerated, 7.5 grams of borneol separating out. This corresponds to a total of 30 per cent borneol. After the separation of the borneol the oil was again fractionated, and the portion above 195° C. yielded, when frozen, an additional 2 grams of borneol, making a total of 9.5 grams, or 38 per cent, of total borneol separated from the oil. The theoretical quantity of borneol in the oil, as shown by the acetylization value, is about 43 per cent, the lower percentage which was actually obtained being caused by incomplete separation due to the smallness of the amount saponified.
Esters of borneol.—A careful examination of Table IV shows that the esters of borneol, possibly chiefly bornyl heptoate and valerianate, are found in the fractions boiling above 190° C., principally in the highest boiling fractions; a perfect separation of these esters was not feasible because of the existence of the esters as mixtures of several acids. The ester numbers of the fractions, however, show the distribution of the esters at the different temperatures.
SUMMARY.
Briefly summarizing the results of the analyses, the oil of wild sage may be said to be composed: (1) Of total borneol camphor, 43 per cent, of which about 6.8 per cent exists as bornyl heptoate (calculating the esters of the oil as heptoic acid salts of borneol), leaving 35.8 per cent of free borneol camphor present in the oil; (2) of cineol (eucalyptol), 18 to 20 per cent; (3) of fenchone, 8 to 10 per cent; (4) of free acids, chiefly œnanthylic, or heptoic, acid, 0.58 per cent, with traces of formic and caprylic acids; (5) of combined acids in form of esters, chiefly, œnanthylic acid, with smaller quantities of valerianic, undecylic, and formic acids. It is very probable that a small amount of terpenes were also present in the portion distilled below 175° C., which, however, were not identified.
As will be noted from the above, the chief constituents of the oil of wild sage are borneol camphor and cineol, each of which possesses valuable antiseptic qualities. Since there is a high percentage of these constituents, the oil from this wild plant should prove of value for medicinal purposes. Another important use of the oil is suggested by the high content of borneol, a constituent which finds application in celluloid manufacture, and which is readily separated from this oil. Lastly, combining the agreeable aromatic quality with its antiseptic qualities, the oil should prove important as an ingredient of medicinal soaps or as a scenting substance.
Inasmuch as the wild sage plant grows chiefly on sandy and stony hills which are practically waste lands and which require but little moisture, it would seem that the plant could be cultivated in various sections of the Northwestern States.
SWAMP BAY.
BOTANICAL DESCRIPTION AND DISTRIBUTION.
Persea pubescens (Pursh.) Sarg., commonly known as swamp red bay or swamp bay (figs. 5 and 6), is an aromatic evergreen tree attaining a height of 30 feet or more, but usually occurring as a shrub. The leaves and twigs of the tree possess a pleasant camphoraceous odor. The swamp bay occurs abundantly in swamps and hammocks from North Carolina to Florida and Texas. The tree is a member of the family Lauraceæ, to which the camphor tree belongs.
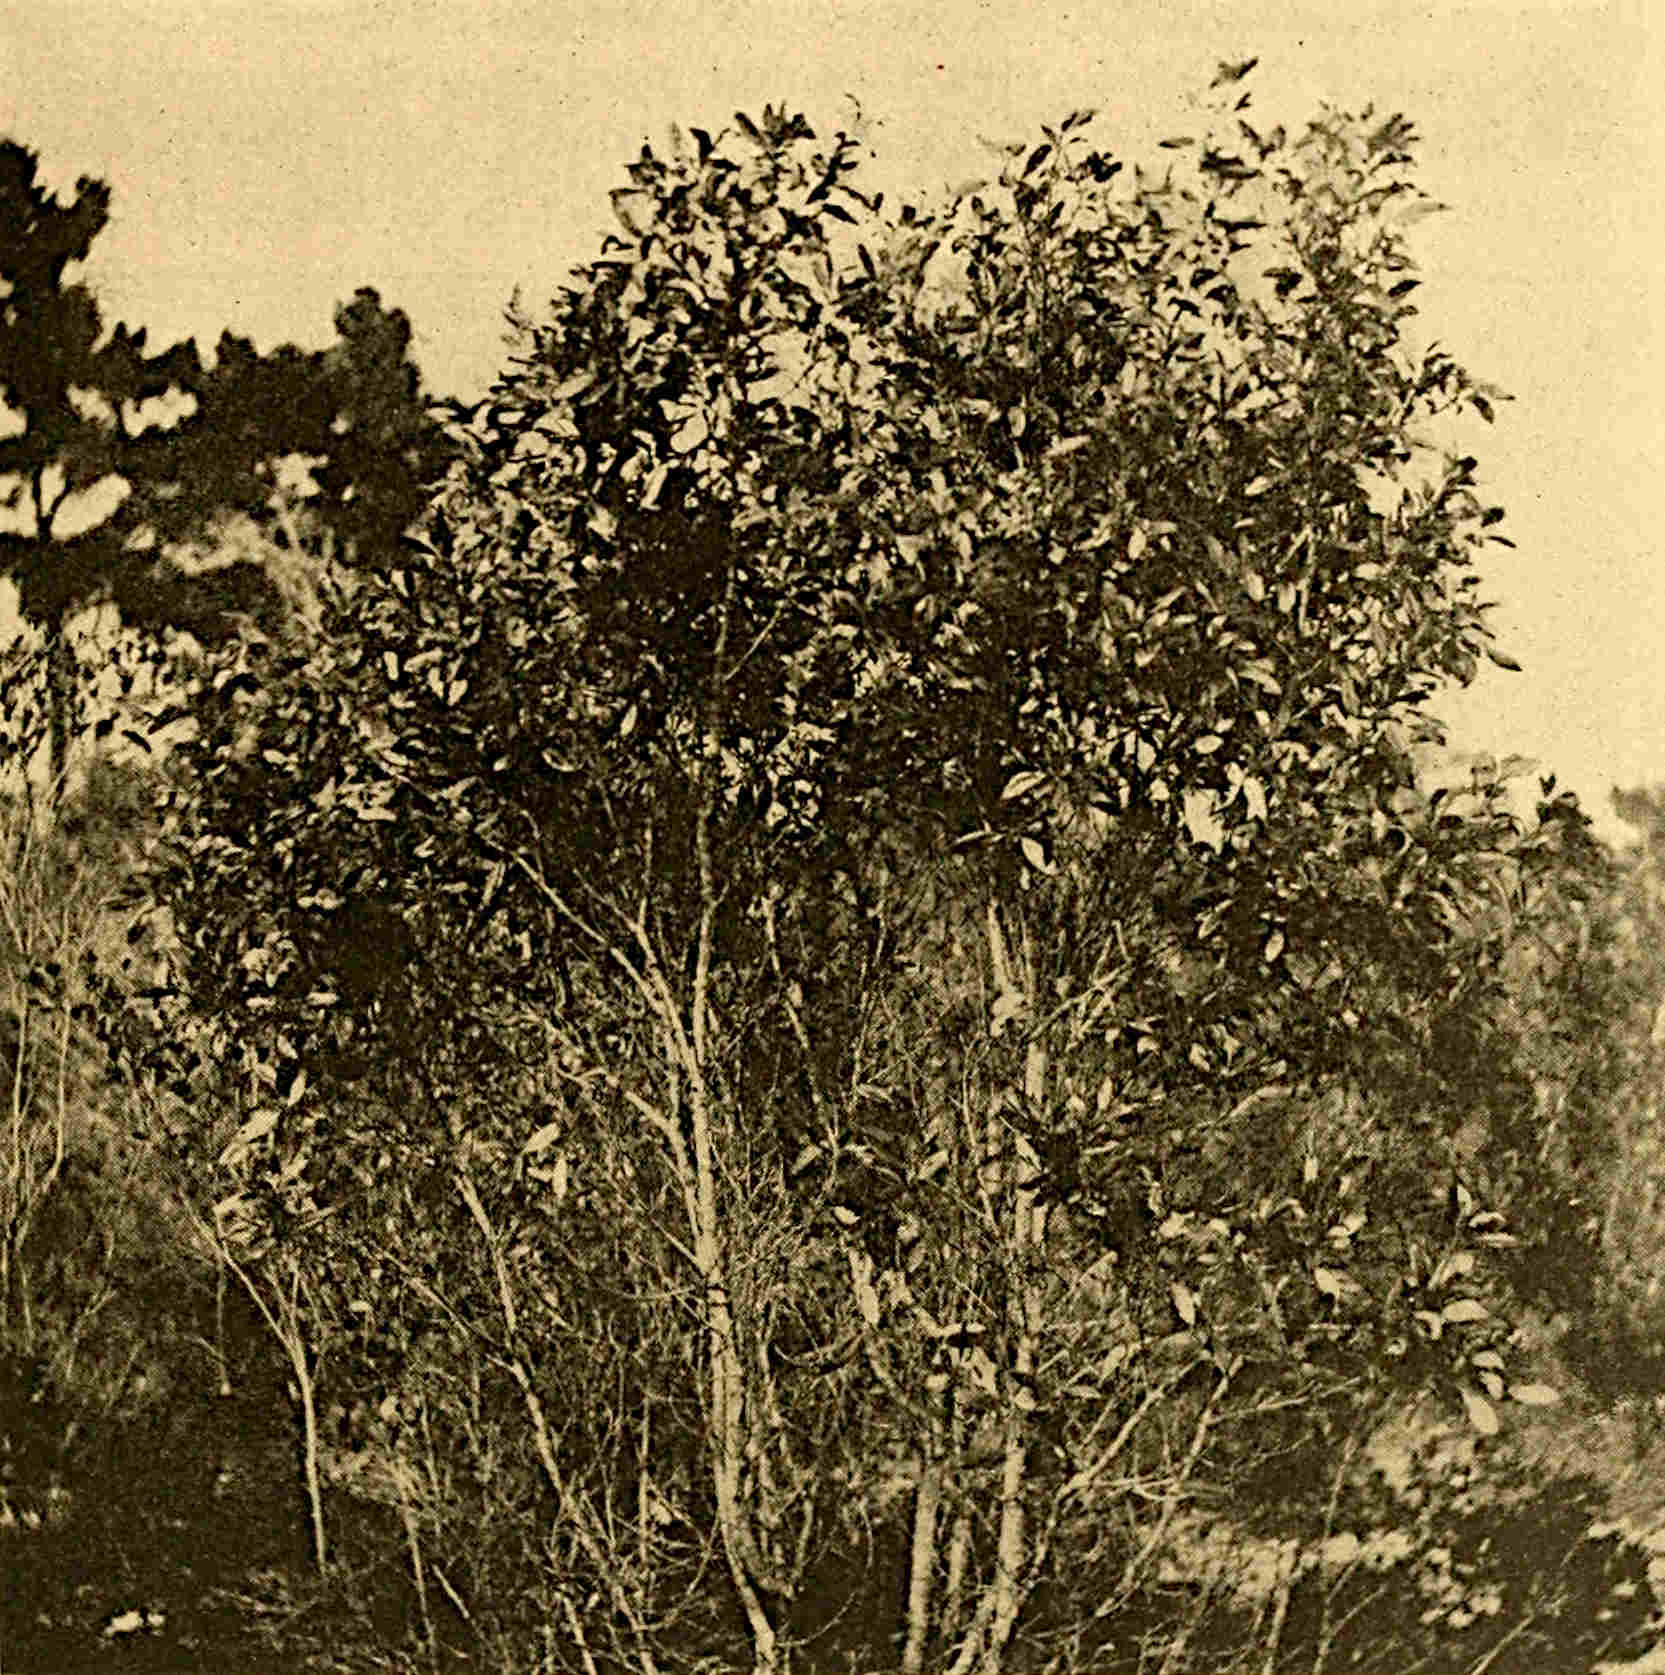
Fig. 5.—A swamp bay tree (Persea pubescens) growing near Orange City, Fla.
DISTILLATION OF THE OIL.
Because of the strong camphoraceous odor and its close relationship to the camphor tree, the extraction and possible utilization of the oil from this wild aromatic plant suggested itself. Accordingly, during the summer of 1910, with the assistance of Mr. S. C. Hood, in charge of the station at Orange City, Fla., a small quantity of the leaves and twigs of this plant was distilled and a yield of about 0.2 per cent of oil was obtained. But with proper conditions and precautions the yield could no doubt be very materially increased, depending largely upon the time at which the distillation is made, and also upon the proportion of twigs and branches included. The above distillation was made late in the summer, long after the blossoming period, the stage at which a plant is usually most productive in volatile oils, and the material also contained many branches and much woody matter.
The oil obtained was pale yellowish brown in color, with a strongly aromatic and camphoraceous odor, and a persistent bitter, slightly pungent, and camphorlike taste. The specific gravity at 25° C. was 0.9272; specific rotation, AD = +22.4°; refraction, ND 25° = 1.4695. The oil was soluble in one-third its volume of 80 per cent alcohol, becoming faintly turbid upon the addition of five volumes or more of alcohol.
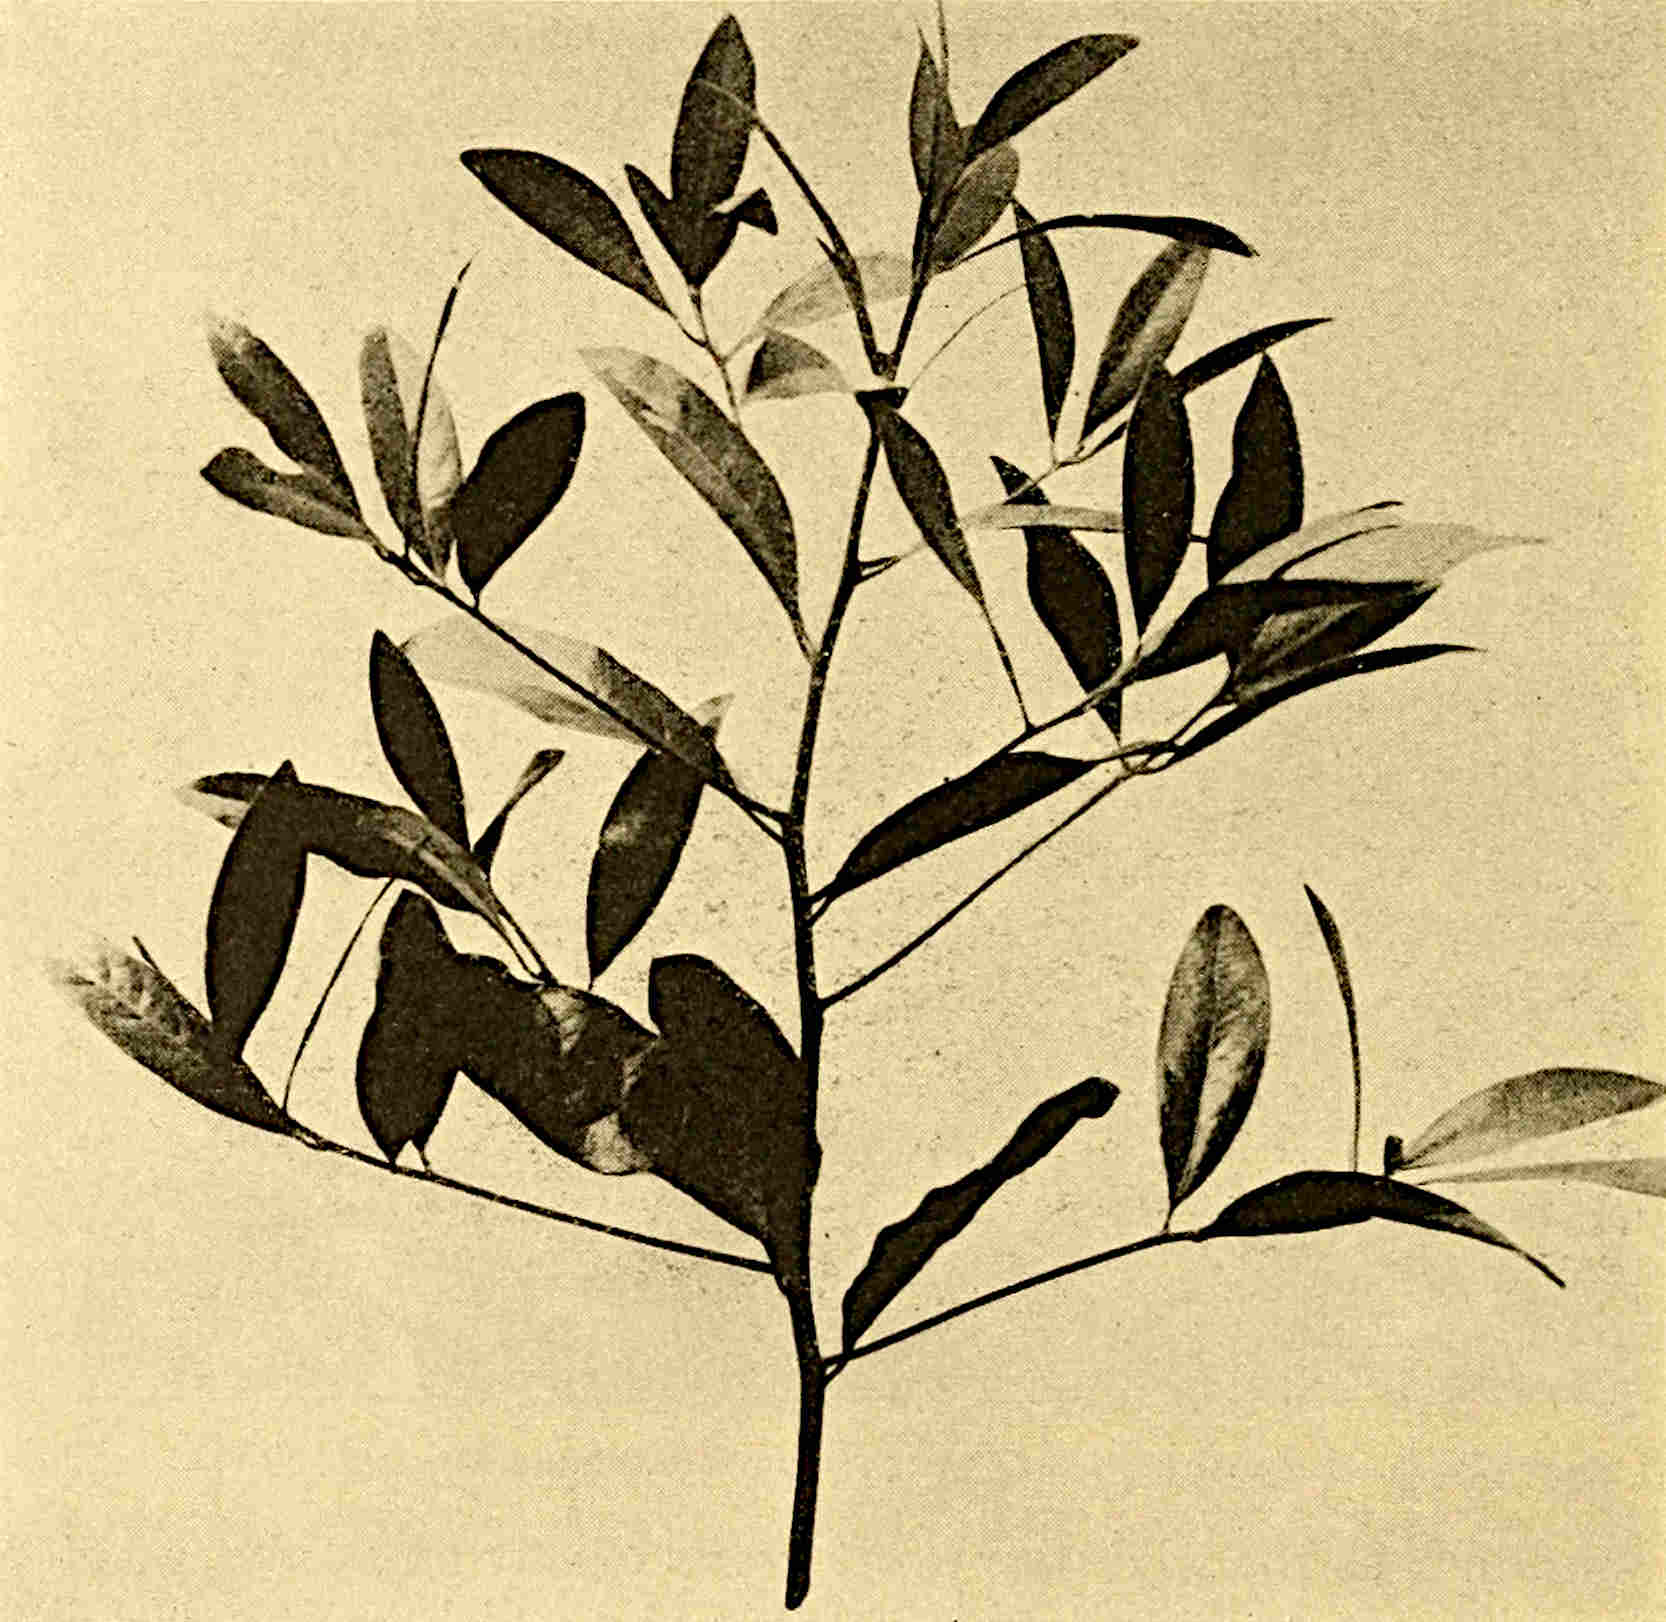
Fig. 6.—A small branch of swamp bay.
CHEMICAL EXAMINATION OF THE OIL.
CHEMICAL CONSTANTS.
A preliminary examination of the oil disclosed considerable free acidity, the acid number being 2.8, while the ester content was rather low, the ester number being 14.5. The low ester number would seem to indicate a low percentage of alcoholic compounds in combination with acids, and would correspond to 4.9 per cent of esters calculated as the acetate of borneol. After acetylization of the oil with acetic anhydrid the saponification number was found to be 64, which corresponds to 14.6 per cent of free alcohol, calculated as borneol.
In order to identify conclusively the constituents of the oil and the forms in which they occur, and to separate quantitatively the predominant constituents, the oil was subjected to a more careful and detailed analysis.
FREE ACIDS.
The free acidity of the oil as indicated by the preliminary tests was removed by shaking with 10 per cent aqueous sodium carbonate solution in several portions. The aqueous alkaline extracts, after being deprived of any adhering oil by extraction with ether, were concentrated, acidified, and distilled with a current of steam. The acids which were obtained separated principally as oily globules on the aqueous distillate, which was only faintly acid.
The free insoluble acids which were separated from the aqueous distillate by extraction with ether and evaporation of the solvent were neutralized with a solution of potassium hydroxid and then precipitated in fractions with a solution of silver nitrate.
Fraction 1. 0.0227 gram silver salt gave 0.0130 gram silver = 57.2 per cent silver.
Fraction 2. 0.0213 gram silver salt gave 0.0119 gram silver = 55.8 per cent silver.
It appears from the above results that the only acid existing in the free state in the oil is butyric acid, since silver butyrate gives theoretically 55.3 per cent of silver, fraction 1 being slightly contaminated, due possibly to a slight excess of potassium hydrate which was added when the acids were neutralized and which would appear in the first precipitate.
From the remaining faintly acid distillate, after neutralization with barium carbonate and concentrating, only a trace of precipitate, insufficient for silver determination, resulted upon the addition of silver nitrate solution. The butyric acid detected in the free insoluble acids was evidently extracted by the ether, in which it is very soluble.
COMBINED ACIDS.
As stated previously, the oil was found to contain a small percentage of esters, or organic acids in combination with higher alcohols. In order to identify these acids, which are in combination in the form of esters, a quantity of the oil, after removing the free acids, was saponified by heating on a water bath for half an hour with a slight excess of alcoholic potassium hydroxid. The mixture, after saponification, was diluted with water and the unsaponified oil separated. The alkaline liquid, which now contained the combined acids as their potassium salts, after being freed from adhering particles of oil by shaking with ether, was acidified with sulphuric acid and distilled with steam. The insoluble oily acids which formed on the distillate were separated by shaking the distillate lightly with ether and evaporating the ether.
SOLUBLE COMBINED ACIDS.
The aqueous portion of the distillate which contained the soluble combined acids of the oil was neutralized with barium carbonate, concentrated and precipitated with silver nitrate solution. Only a small precipitate resulted. This precipitate was found to contain 55.9 per cent of silver, which corresponds to silver butyrate. Hence the acid in the distillate was butyric acid.
INSOLUBLE COMBINED ACIDS.
As heretofore stated, the insoluble oily acids obtained by extraction with ether were carefully neutralized with potassium hydroxid solution and precipitated fractionally with silver nitrate. Two precipitates were obtained which were thoroughly washed and dried. The first and largest precipitate assayed 51.2 per cent silver, the second assaying 45.1 per cent silver. This would indicate that the insoluble acids were valerianic acid (silver valerianate requiring 51.6 per cent silver), and heptoic acid (silver heptoate requiring 45.5 per cent silver), the valerianic acid predominating.
The results show that the esters of this oil exist as the salts of butyric, valerianic, and heptoic acids, valerianic acid esters, however, predominating.
FRACTIONATION OF THE OIL AND SEPARATION OF THE STEAROPTENE.
For the purpose of accomplishing a separation of the constituents, 50 grams of the oil, after saponification, were dried and subjected to fractional distillation in a three-bulb Ladenburg flask. The results are given in Table V.
Table V.—Fractionation of saponified oil of swamp bay and description of fractions.
Fraction. Temperature. Distilled. Remarks.
Degrees C. Per cent.
1 Below 170 1.1 Penetrating odor; largest portion of the fraction distilled over below 80° C.; temperature rose rapidly to 170° C.
2 170 to 182 8.8 Camphoraceous cineol-like odor; largest portion distilled 175° to 180°.
3 182 to 185 9.2 Strong cineol-like odor; temperature rose uniformly.
4 185 to 190 13.5 Cineol-like camphoraceous odor; temperature rose uniformly.
5 190 to 195 13.0 Strong camphoraceous odor; temperature rose uniformly.
6 195 to 200 5.8 Strong camphorlike odor; crystals appeared in condenser;[34] largest portion distilled between 198° to 200° C.
7 200 to 205 12.5 Strong camphorlike odor; fraction semisolid upon cooling; temperature rose uniformly.
8 205 to 215 14.0 Strong camphorlike odor; fraction almost solid upon cooling; distilled largely between 205° to 210° C.
9 215 to 225 12.5 Strong camphoraceous odor; fraction semisolid; temperature rose uniformly.
10 225 and above 9.0 Heavy yellow oil with camphoraceous odor.
Beginning with fraction 6 each successive fraction was refrigerated in a freezing mixture of ice and salt and the crystals separated by centrifuging in a platinum Gooch crucible. A total of 13.7 per cent of crystals was obtained.
In order to obtain a further separation of crystals the portions of the oil beginning with fraction 5 were fractionated into the following fractions: 190° to 195° C.; 195° to 200° C.; 200° to 205° C.; 205° to 215° C.; 215° to 233° C.; 233° to 260° C. A total of 4 per cent of crystals was obtained by refrigeration and centrifugation of those fractions in which crystals appeared. The portion between 190° and 215° C., and also fraction 4 of the original, were further fractionated into four parts: 185° to 190° C.; 190° to 195° C.; 195° to 205° C.; 205° to 215° C., an additional yield of 3.3 per cent of crystals being obtained.
By the above method of successive fractionation and refrigeration a total of 21 per cent of crystals was obtained from the oil. This represents only approximately the total percentage of stearoptene in the oil. The separation was not at all quantitative, as a considerable proportion was lost in the manipulations incident to the separation. Since the quantity of oil at hand was so meager the fractions were reduced to such small quantities that further separation of crystals was impossible, and as unavoidable losses were encountered in transferring to and from the centrifuge the final percentages were materially affected and the true amount of stearoptene may be assumed to be considerably more than is shown above.
After the fractionation and refractionation of the oil and the separation of the stearoptene portion, the remaining elaoptene portion grouped itself into fractions, whose physical properties were determined and qualitative tests for their constituents applied, as shown in Table VI.
Table VI.—Refractionation of the oil of swamp bay, showing the physical properties of the fractions.
Fraction. Temperature. Specific gravity at 25° C. Rotation in 50 mm. tube. Re-fraction ND 25°. Tests applied.
Degrees C. Degrees.
1 Below 170 Insufficient. Insufficient. 1.4648 When shaken with water the aqueous solution strongly reduced magenta solution to violet color; also produced silver mirror with ammoniacal silver nitrate.
2 170 to 182 0.9011 +22.5 1.4630 Iodol (tetraiodopyrol) dissolved in oil by gentle warming yielded yellow crystals melting at 115° C.; cineol iodol melts at 112° C.
3 182 to 185 .9012 +21.5 1.4628 Treated with iodol and the yellow crystals recrystallized from benzol melted sharply at 112°.
4 185 to 190 .9075 +23 1.4628 Cineol-iodol crystals melted at 113° C.
5 190 to 205 .9228 +31 1.4653 Do.
6 205 to 215 .9351 1.4706 Negative test with iodol.
7 215 to 233 .9358 1.4765 Do.
8 233 to 260 .9360 1.4830 Oxidized with 3 per cent potassium permanganate in cold yielded camphor crystals.
IDENTIFICATION OF THE CONSTITUENTS OF THE OIL.
Camphor.—The compound obtained from the oil by refrigeration was a soft, white, granular, crystalline mass, and possessed a distinct camphorlike odor and slightly bitter camphoraceous taste. The crystals sublimed readily and melted at 174° to 176° C. The boiling point of the compound was 205° C., and the rotation in a 50 mm. tube of 20 per cent solution in alcohol was found to be +3.8°, 20 percent solution of commercial camphor in alcohol rotating +3.5°. It was readily soluble in alcohol and the other organic solvents.
To further identify the crystals with ordinary camphor two compounds were prepared, the semicarbazone and the oxime, with which camphor forms definite chemical compounds. The semicarbazone was prepared according to the method of Tiemann. (See p. 17.) The crystals obtained after recrystallization from alcohol melted at 237° to 239° C., pure camphor semicarbazone melting at 236° to 238°. For the preparation of the oxime Auwers’ method was applied. (See p. 16.) Recrystallized from ether the oxime melted at 117° to 118° C., whereas pure camphor oxime melts at 118° to 119° C.
Since the physical and chemical properties of this substance correspond almost identically with those of camphor, it may be safely stated that the crystals are those of commercial dextro camphor.
Aldehyde constituent.—From the pungent and penetrating odor and the strong reducing properties of the first fraction, which, as shown in Table V, distilled largely below 80° C., there would seem to be the possible presence of a trace of formaldehyde.
Cineol, or eucalyptol.—Qualitative tests as indicated in Table V show the presence of cineol in fractions from 170° to 205° C., the characteristic crystalline cineol addition product of iodol corresponding in melting point to the pure cineol iodol. Cineol was further identified in these fractions by the preparation of cineol hydrobromid prepared by passing dry hydrobromic acid gas into a well-cooled solution of the oil in petroleum ether. A crystalline hydrobromid was obtained from each fraction which gave the iodol reaction. The hydrobromids prepared melted between 55° to 57° C., while pure cineol hydrobromid is reported as melting at 56° to 57° C.
Since the presence of cineol in the several fractions of the oil was proved, a quantitative estimation was deemed desirable. Because of the smallness of the individual fractions the hydrobromic acid method was adopted in this estimation, it being the most accurate when cineol is present in only small quantities. The phosphoric acid method is best adapted to oils which are very rich in the compound. The hydrobromic acid method has been used in the assay of eucalyptus oils,[35] and consists essentially in conducting dry hydrobromic acid gas into a solution of the oil in about twice its volume of petroleum ether, the solution being well cooled by a freezing mixture, separating the crystals on a force filter, washing and decomposing with water, and measuring the cineol formed. A slight deviation was made from the directions on account of the smallness of the fractions and consequently the small amount of hydrobromid obtained, which when decomposed with water would introduce an error. After the hydrobromid of cineol was obtained in each case and washed it was weighed and the percentage of cineol was calculated from the weight of the crystals from a given quantity of each fraction. In this manner by assaying the four fractions which gave qualitative tests there was found to be a total of 19.8 per cent of cineol in the oil.
Borneol.—By oxidation of fraction 233° to 260° C. with a 3 per cent solution of potassium permanganate, slightly warming and allowing it to stand for 12 hours, then shaking out the mixture with ether and allowing the ether to evaporate, a mass of crystals remained which proved to be camphor. It is possible that borneol was present in this fraction, as borneol is readily oxidized to camphor with ordinary oxidizing agents. Since the preliminary chemical examination of the oil indicated a small percentage of esters and of free alcohol, the alcohol was probably borneol.
SUMMARY.
From the results obtained in the chemical examination it appears that the oil of swamp bay contains over 21 per cent of camphor, 19.8 per cent cineol, and borneol, the latter possibly occurring to a small extent as esters and as the free alcohol. No terpenes were identified. Since only a very small portion of the oil distills over below 175° C., it would seem that the oil is not terpenic in character, as most members of the terpene group of hydrocarbons boil below 175° C.
Besides the constituents mentioned, the oil contains butyric acid in free condition to a slight extent; butyric, valerianic, and heptoic acids combined in the oil as esters, valerianic acid predominating, and a slight trace of an aldehyde, possibly formaldehyde.
This oil possessing, as has been proved, considerable quantities of such constituents as camphor, cineol, and borneol, all of which are valuable therapeutic agents, may be of economic importance from the standpoint of the perfumer or the medical practitioner. Doubtless if the distillation of the plant were carried on, attention being paid to the stage of growth at which it is distilled and the distillation restricted to the leaves and small twigs, the yield of oil and possibly the yield of the three important constituents mentioned could be considerably augmented.